+86 13638632902
 Jason@cleanroomequips.com


 MENU
The first portion of the SOP will cover the next steps in the process of sedimentation bacteria testing, the regulatory requirements, and the use of both active and passive methods.
The methods will be discussed so that the readers will be informed about what techniques to use as well as what techniques will be taught.
The next section will cover the basic principles of the sedimentation bacteria testing process, which includes how gravity plays an important role in the process and what types of particles can be used.
After discussing the principles of the sedimentation bacteria testing process, the next area will cover the use of active versus passive methods of air sampling, forming the base of any cleanroom microbial monitoring SOP.
To conclude the discussion of sedimentation bacteria testing, the reader will be made aware of the procedures to follow in order to perform rules set forth by the pharmaceutical industry to maintain compliance with the standards and guidelines.
In order to understand this SOP, the reader must understand two important sections of this protocol.
The first stage in this procedure will cover the fundamentals that will be helpful in sounding decisions as well as making appropriate rules.
Once again, the reader will now be taught about sedimentation bacteria testing because it is important that they understand how each of these procedures works.
Fundamentals of Sedimentation Bacteria Testing
There are many ways to understand how gravity comes into play when performing the sedimentation bacteria test.
Gravity does play an important role in the sedimentation bacteria testing procedure because there is no gravity acting upon airborne microorganisms.
The particles that are greater than five micrometres will settle at a given rate during air sampling.
Principles of Settle Plate Method cleanroom monitoring
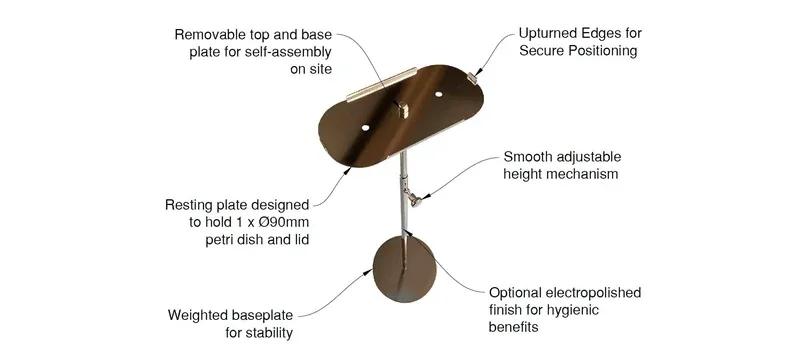
The settle plate method cleanroom approach has been used for over a century. Koch's principle established that colony count reflects the surrounding air's microbial burden.
Nowadays, pharmaceutical purification laboratories apply controlled exposure times and standardized 90 mm plates in order to get valid results and collect field studies for recovery. The enumeration of colony-forming units gives air quality a semi-quantitative value.
The method does not give the volumetric concentration immediately. The findings will usually present CFU per plate for every period of exposure, mostly expressed as CFU per 4 hours.
Many parameters influence the depletion rate, including: air speed, temperature and humidity, as well as the size distribution of the particles. Turbulent air flow near the air diffuser or door makes particles stay in the air more.
The method does not provide information about the concentration of particles smaller than 1 µm, since they stay in the air for the longer time. Information received in this way is semi-quantitative and specific to the particular site.
Active vs Passive Air Sampling Comparison
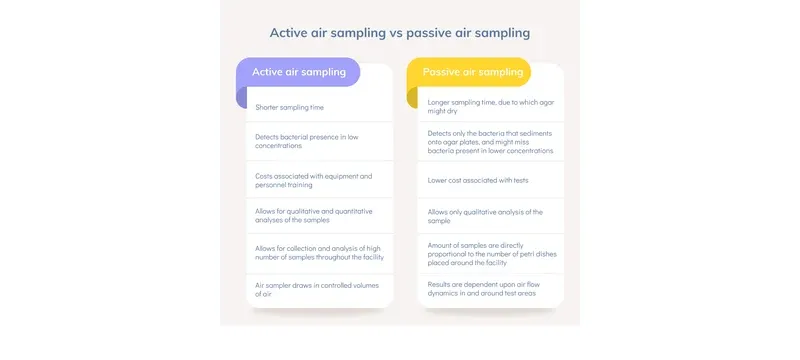
Active sampling uses known air volumes through either impaction or centrifugation. Passive sampling relies on gravity. This active vs passive air sampling comparison underpins pharmaceutical microbiology testing in every graded zone.
Table 1: Comparison of Active and Passive Air Sampling Methods
| Parameter | Active Sampling Technique | Passive Settle Plate Sampling Technique |
|---|---|---|
| Methodology | Forced Impaction or Centrifugation | Gravitational Settling |
| Result Units | CFU/m³ | CFU/plate/4 hours |
| Air Volume | Fixed and Controlled | Not Fixed |
| Impulse Flow Disruption | Possible at Extremely High Flow rates | Negligible |
| Legal Limitations | Required in Grade A and B of GMP (EU Annex 1 [6]) | Required in Grade A and B of GMP (EU Annex 1 [6]) |
| Ideal Application | Volumetric trend measurements, critical zones | Surveillance at constant low cost |
| Parameter | Active | Passive |
|---|---|---|
| Methodology | Forced Impaction | Gravitational Settling |
| Result Units | CFU/m³ | CFU/plate/4 h |
| Air Volume | Fixed | Not Fixed |
| Flow Disruption | Possible | Negligible |
| Legal Limits | Grade A and B [6] | Grade A and B [6] |
| Ideal Use | Critical zones | Low cost surveillance |
Active sampling measures volumetric concentration, whereas passive plates monitor around multiple locations inexpensively. The GMP also mandates the use of the active and passive sampling methods in Grade A and B areas.
Regulatory Framework for GMP Environmental Monitoring
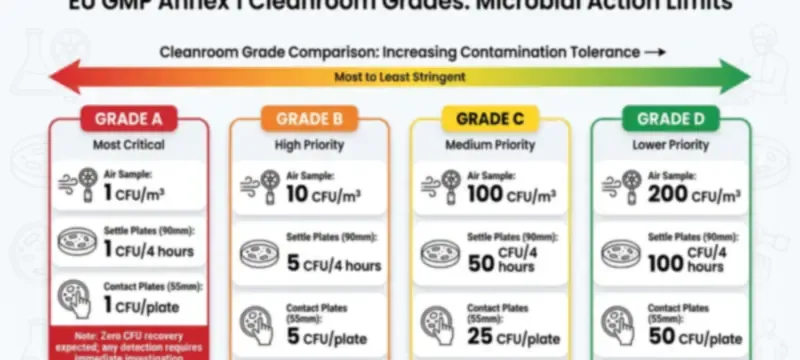
The GMP legislation states the highest permissible contamination levels for viable materials in the grades A, B, C, and D [6]. Section 21 CFR 210/211 also mentions cleanliness and environmental control practices [9].
Section 211.113 of the FDA also states that there have to be written procedures in place for environmental monitoring of the sterile production area [9]. iso 14644-1 defines the classification of cleanliness of particles [1]. As per iso 14644-2, monitoring and requalification have to be carried out [2].
USP section <797> and <1116> provides guidance for monitoring of microorganisms in sterile formulation and controlled environment [3][4]. PIC/S GMP guide annex 1 is similar to that of Europe [7]. GB 50073 and GB 51110 are based on the standard ISO classifications and the minimum monitoring requirements in pharmaceutical facilities in China [10][11].
Regulatory Compliance Framework - Outlined Requirements
When a suitable regulatory compliance framework is developed, it will reflect the relevant regulations that need to be adhered to. There are critical requirements that have to be taken into consideration as shown below:
- The time plates should have a maximum of four hours of exposure in Grade A and B cleanroom environments, while in grade c and D cleanrooms, the exposure time is determined through quality risk management.
- Continuous monitoring must occur in the Grade A areas for verification of microbial growth in the cleanroom environment.
- Should there be any presence of microbial growth in the Grade A cleanroom areas, a deviation report must be prepared.
- The position for the monitoring must come from proper risk assessment.
Cleanroom Microbial Monitoring SOP Framework
The cleanroom environmental monitoring standard operating procedures (SOP) must include the relevant purpose, scope of monitoring, responsibilities, frequency, position of monitoring and the documentation.
The SOP must also be periodically reviewed on an annual basis and may be revised following any significant changes in the processes or in the cleanroom facility.
SOP Structure and Documentation Requirements
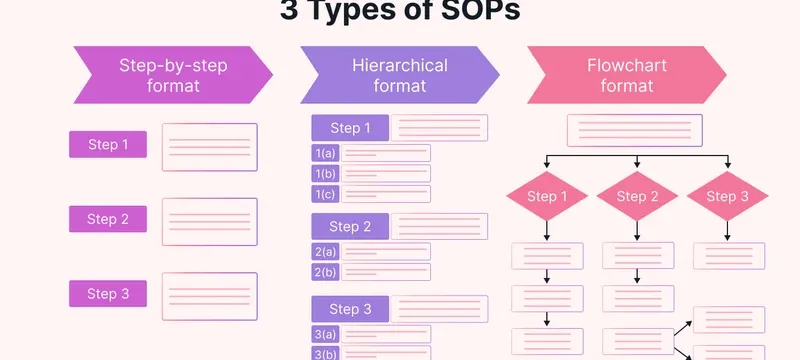
The cleanroom SOP has to include the statement of purpose and scope at the beginning. The current standard references should include the full name of the relevant references and standards that were followed.
The prescribed standard references should include the following: EU GMP Annex 1, ISO 14644-1/-2, and the United States Pharmacopoeia (USP) <1116>.
All equipment, materials and media used must be specified in the SOP. There must be explicit instructions on the step-by-step procedures. The relevant equipment and materials include Tryptic Soy Agar (TSA), plates, and labels.
Table 6: SOP Essential Elements
| Element | Requirement |
|---|---|
| Purpose and scope | Clear definition of programme boundaries |
| Normative references | Full standard names with edition dates |
| Equipment specs | Media type, plate size, incubator parameters |
| Sampling procedure | Step-by-step numbered instructions |
| Incubation protocol | Temperature, duration, counting rules |
| Reporting format | Standardised data entry template |
| Deviation handling | Investigation and escalation workflow |
| Revision control | Version history and approval signatures |
| Element | Requirement |
|---|---|
| Purpose and scope | Programme boundaries defined |
| Normative references | Full names with edition dates |
| Equipment specs | Media, plate size, incubator |
| Sampling procedure | Numbered step-by-step instructions |
| Incubation protocol | Temperature, duration, counting |
| Reporting format | Standardised data template |
| Deviation handling | Investigation workflow |
| Revision control | Version history and approvals |
Sampling Frequency and Location Planning
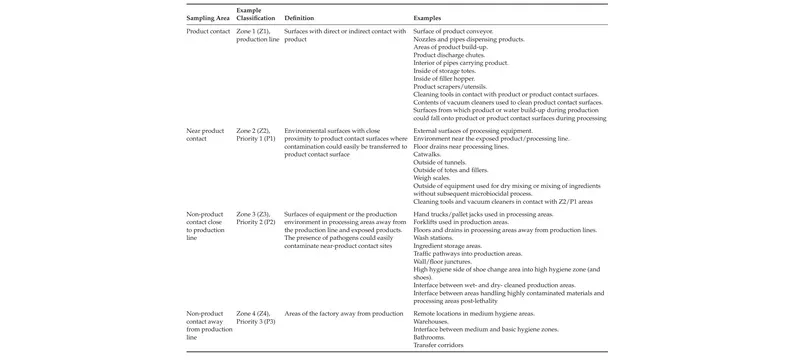
The intensity of monitoring in iso 5 zones must be the highest whereas in iso 8 zones, lower frequency monitoring can be accepted. When it comes to sampling, the critical zones where open products are present must be considered.
Traffic of personnel leads to deposition hence, those zones where a high number of people visit must be monitored more.
The placement of Deiiang™ terminal HEPA filters and sampling sites must be taken into consideration for optimum productivity given the fact that the plates on the filtered side would give different results compared to those in the corners.
Jason Peng is a cleanroom designer working for Deiiang. He highlights the fact that plates placed directly under HEPA filters will have lower counts (30-40%) because the downward velocity of the air will lead to sweeping of particles away from the agar.
Table 2: Recommended Sampling Frequency by cleanroom grade
| Grade of Cleanroom | ISO Class | Frequency of Sampling (Settle Plates) | Typical Locations for any Shift |
|---|---|---|---|
| Grade A | ISO 5 | Continuous while operations | 2-4 critical points |
| Grade B | ISO 5 | Once in a Shift | 4-6 points |
| Grade C | ISO 7 | 2-3 times a week | 4-8 points |
| Grade D | ISO 8 | Once a week | 4-6 points |
| Grade | ISO Class | Sampling Frequency |
|---|---|---|
| Grade A | ISO 5 | Continuous while operations |
| Grade B | ISO 5 | Once in a Shift |
| Grade C | ISO 7 | 2-3 times a week |
| Grade D | ISO 8 | Once a week |
To illustrate, a grade c cleanroom that is monitored 3 times a week would amount to 18 plates a week and 936 plates in a year.
Roles and Responsibilities for Execution
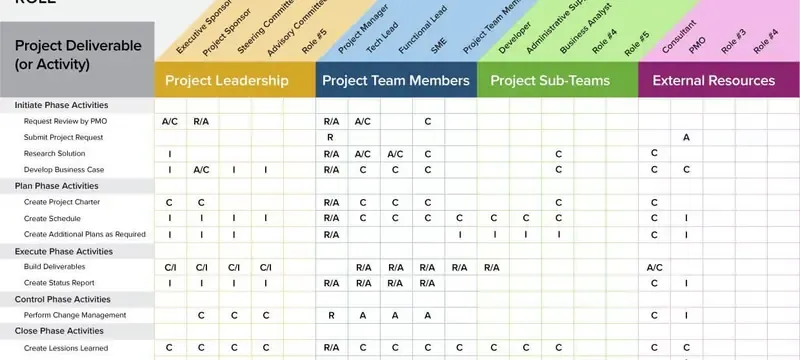
QA reviews trends and gives corrective actions once alert level or action levels are reached. Quality Control is responsible for sampling, incubation, counting, and reporting on time.
Engineering department of the facility is responsible for the HVAC system and also documents filter changes, and keeps an air change rate record. The validation team does reproducible tests to qualify the SOP and to document.
Step-by-Step Sedimentation Bacteria Testing Procedure
During this stage, it will be necessary to ensure that each step is documented. In this section, we will cover the process of preparing, sampling in the field, incubating, and calculating the results.
Pre-Testing Preparation and Media Selection
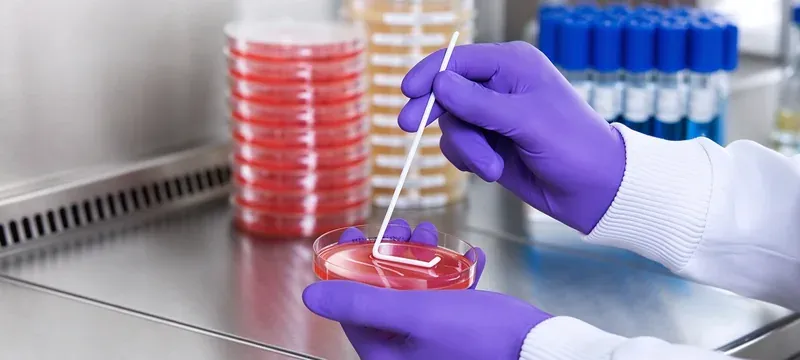
The most widely used medium for bacterial recovery is tryptic soy agar (TSA) & for each batch of TSA that is prepared, it will be necessary to validate the sterility of the media with verification. It will be necessary to label each of the TSA plates with a batch identification code, date, and operator identification.
Plates that are intended for use in the sampling process should be transported within sealed containers that allows the temperature of the media to remain stable so that there is little to no condensation on the media or other contamination issues that may arise otherwise. In the case of unused plates, they should be kept at a temperature of 2-8 degrees Celsius until they are needed.
In addition, each employee must properly gown themselves in the validated method prior to entering the pharmaceutical purification cleanroom prior to sampling.
The preparation activities are as follows:
Sampling Execution in Pharmaceutical Purification Workshop
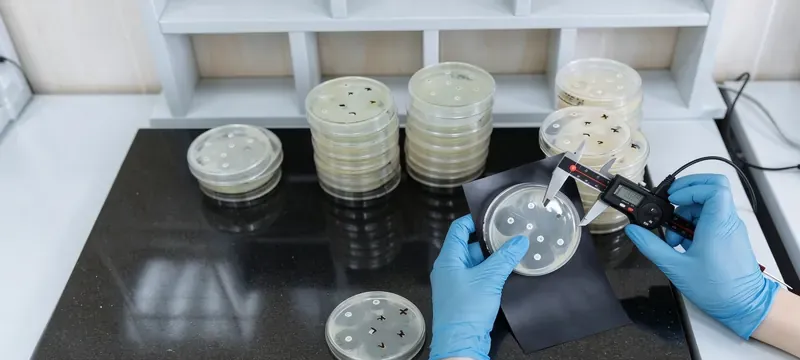
The height at which you should place your plates should be approximately 1 meter off the ground or at working height. The lid of the plate is then lifted to expose the agar to contaminants while ensuring that no more than 4 hours will pass during which it is exposed.
The doors must remain closed while sampling in order to prevent any turbulence or air disruption from the outside world. The person sampling should also try and remain at least 1 meter from the plates in order to avoid any shedding of contaminants.
There should be duplicate plates for each sampling event to help determine the variations in counts over time. You will also want to include negative controls in order to ensure that the media used is not contaminated.
When exposing the agar plates, if there is any door opening that occurs more than 10 seconds or any person enters the area within 1 meter of the exposed plates, then the exposure process must start back over again with a new plate. Record all interruption in the sampling log, including time and reason for the interruption.
Checklist for Sampling Process
Incubation, Colony Counting and Result Calculation
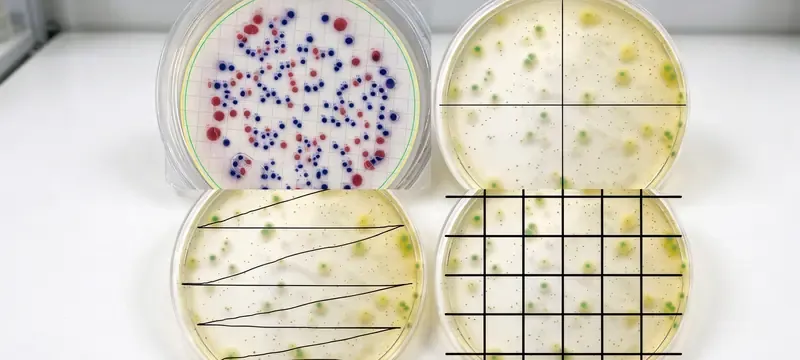
Incubation of exposed plates should take place inverted at a temperature between 30 and 35 degrees Celsius for up to five days and at 20 and 25 degrees Celsius for up to seven days to capture the fungi and yeast.
Count the colony counts when there is enough light and with magnification if necessary. Report the results as the number of colony-forming units (CFUs) per agar plate for the exposure duration.
Counting Rules
- Count all colonies. If there are too many colonies, report as "too many to count" greater than 300 colonies per agar plate.
- Mark and disregard swarming colonies since it prevents the visualization of the entire colony growth.
- Perform a positive control plate with the known bacteria strain every month.
Calculating the CFU/m²
The formula to calculate CFU/m² from the plate area and exposure time, for a 90 mm plate area is π * (0.045²) = 0.00636 m². Thus CFU/m² is given by the formula CFU per plate / 0.00636 m².
Table 5: CFU Calculation Example (90 mm Diameter Plate)
| Input Scenario | Calculation Step | Result |
|---|---|---|
| 12 CFU, 4 h exposure | 12 ÷ 0.00636 m² | 1,887 CFU/m² |
| 6 CFU, 2 h exposure | (6 ÷ 2 h) × 4 h ÷ 0.00636 m² | 1,887 CFU/m² |
| Input Scenario | Result |
|---|---|
| 12 CFU, 4 h exposure | 1,887 CFU/m² |
| 6 CFU, 2 h exposure | 1,887 CFU/m² |
The result calculated represents the deposition that has taken place over that time. Any shorter exposure time could be normalized accordingly. Give average numbers for the duplicates.
Viable Particle Count Pharmaceutical Data Interpretation
Valid interpretations for viable particle count pharmaceutical data should be made with limits and trend analysis in mind. Sedimentation is complementary to active air sampling.
This section will describe how correlating viable particles and sedimentation is done along with what the alert or action levels are per grade.
Correlation Between Sedimentation and Viable Particle Data
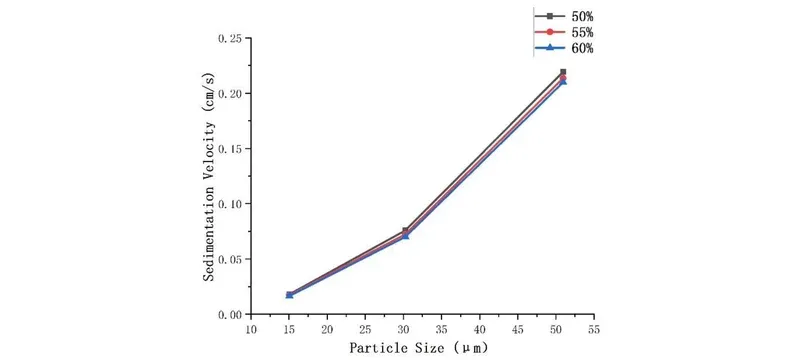
The deposition velocity depends on many factors including particle size, the velocity of the air, and the micro-organisms present. A general approximation that is often used states that a 90 mm settle plate used for four hours will collect culturable colonies equivalent to 1/100th of CFU/m³.
This conversion of 1/100 should be taken with caution and never used in a regulatory submission. Alterations will also occur based upon the speed of air changes and the efficiency of the filter. Fast air changes will not only dilute airborne micro-organisms, but they will also decrease deposition.
Even though the active air count might be the same, the settle plate result may vary between facilities. This fact should always be noted in the SOP and trend report.
Alert and Action Level Setting by Cleanroom Grade
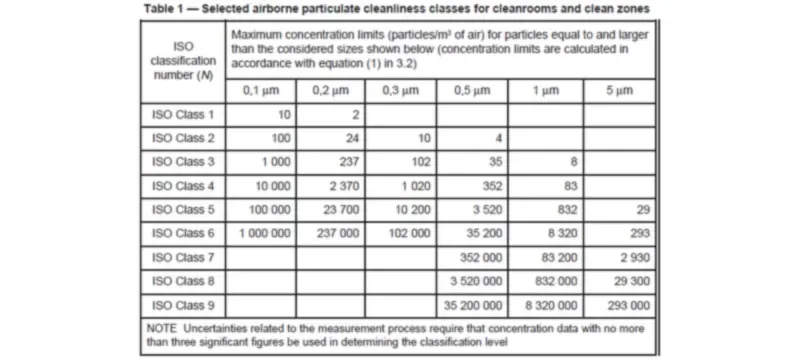
Alert levels indicate a deviation from the norm and trigger additional monitoring. Action levels go beyond acceptable limits and thus require corrective action. The maximum action limits set forth by the EU GMP Annex 1 should be adhered to.
Table 3: EU GMP Annex 1 Viable Contamination Limits and Suggested Internal Alert Levels
| Grade | Action Limit for Settle Plate (CFU/4h) | Action Limit for Air Sample (CFU/m³) | Typical Alert Level (CFU/4h) |
|---|---|---|---|
| A | No growth | No growth | 0 (any growth triggers investigation) |
| B | 5 | 10 | 3 |
| C | 50 | 100 | 25 |
| D | 100 | 200 | 50 |
| Grade | Settle Plate Action Limit (CFU/4h) | Typical Alert Level (CFU/4h) |
|---|---|---|
| A | No growth | 0 (any growth investigated) |
| B | 5 | 3 |
| C | 50 | 25 |
| D | 100 | 50 |
When setting trend adjustments, one must consider previous data along with seasonal variations or process changes taken into account.
For new sites with under 3 months of baseline data, start the initial alert levels by setting at 50% of the regulatory action limit. The alert levels may be modified after 6 months of collecting continuous data.
Cleanroom Biocontamination Control and CAPA
The cleanroom biocontamination control relies on monitoring, investigation, and improvements of the biocontamination process.
When there are out-of-compliance results associated with sedimentation bacteria testing completed, a systematic CAPA process is to be initiated to find out root causes and reach levels that will eliminate future occurrences.
Root cause analysis is included in this subsection.
Root Cause Analysis for Out-of-Specification Results
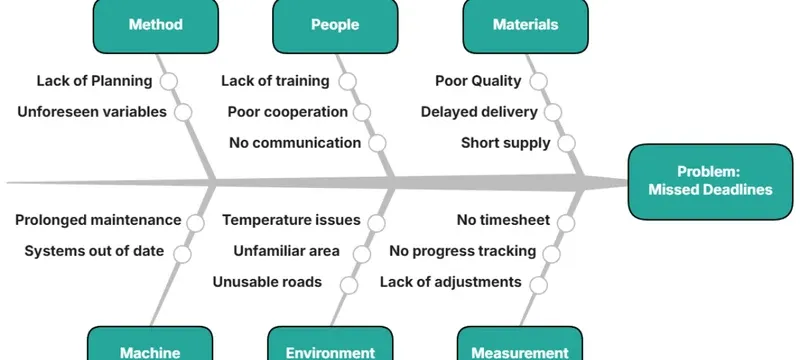
Out-of-specification (OOS) results must be investigated in a systematic manner in four areas: personnel, equipment, facility, and materials. Deiiang data from 28 pharmaceutical cleanroom projects have shown about 62% of sedimentation OOS occurrences are attributed to HEPA seal failure and not the failure of the filter media.
In most cases, the results are due to personnel violations; gowning violations or too much movement into or within the cleanroom result in elevated amounts of deposition.
The HVAC system is evaluated in relation to air change rate, pressure differentials, and filter integrity. DOP or PAO testing is done to show the HEPA has the proper efficiency.
Corrective and Preventive Action Implementation
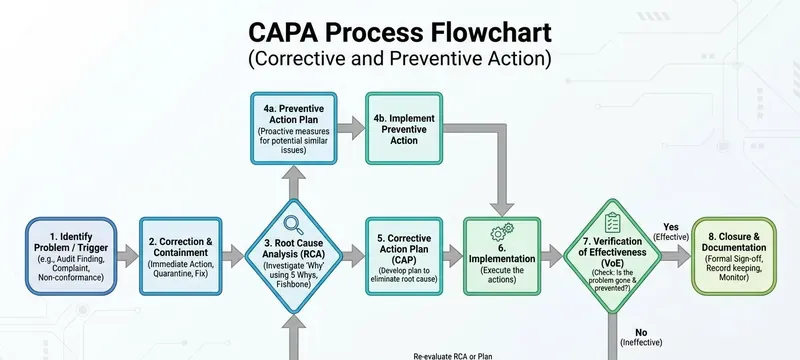
In case of OOS results, an immediate containment action may involve increasing the sampling, increasing restriction to the area, or raising cleaning levels. In general, for the field OOS situation, the root cause must be corrected to bring the product into compliance.
Preventive Actions for Sedimentation Programs
- Conducting HEPA seal integrity checks four times yearly.
- Certifying personnel proper gowning semiannually.
- Changing airflow balances by seasons.
The procedure may entail regular sampling of settle plates for a duration of 14 days following a Grade B violation. New information is incorporated into the SOP.
Deiiang Filtration Solutions for Biocontamination Control
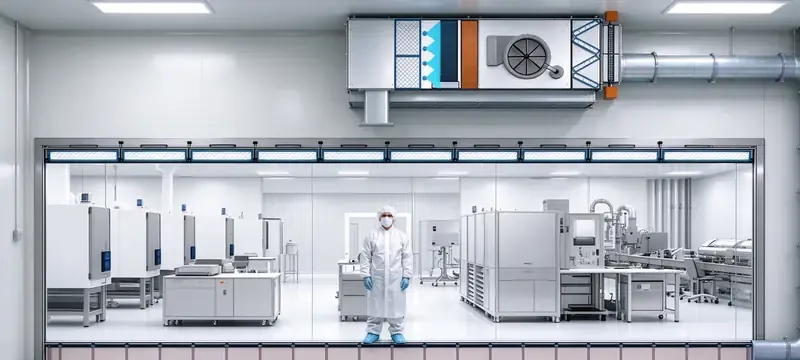
Terminal HEPA filtration serves as a major engineering solution to eliminate airborne microbial contamination. Deiiang™ utilizes high-efficiency filters categorized according to EN 1822 standard ratings as H13 (99.97%-99.99% at 0.3 µm) and H14 (99.995%-99.999% at 0.3 µm).
According to catalogue information, the 592 x 592 x 292 mm size can achieve flow rates of 2,500 m³/h with initial resistance of ≤220 Pa for H14 filters. Using pre-filters such as F7 or F8 bag filters prolongs the life of HEPA filters.
Deiiang™ medium-efficiency glass fiber bag filters have efficiencies of F7 (85%) and F8 (90%) at 0.5 µm according to the old EN 779 standard; ISO 16890 is now utilized as the universal standard, whereas EN 1822 ratings describe HEPA air filter classification schemes.
In a project-reported case for a pharmaceutical packaging cleanroom with an ambition of reaching class 1000 (ISO 6) level and an environment with high levels of ambient particulate matter containing low height plenums, Deiiang™ furnished low profile H14 combined filters (592 x 592 x 292 mm, initial pressure drop of ≤220 Pa and area of 20.04 m², flowrate equal to 2,500 m³/h) that included silicone-free gaskets along with F7 prefilters. DOP testing has been conducted to check the effectiveness on the filter design.
Documented results include the stabilization of particle count to ISO 6 limits starting from ISO 7 levels along with validation that it is possible to save energy.
According to sedimentation testing data, the average CFU/4h count of Grade C areas was decreased from 12 counts before the modification into 3 CFUs after performing the upgrade. Testing and validation provide consistent and accurate outcomes across operator locations.
The parameter of continuous improvement ensures a program that is continuously compliant with the dynamics of its operation environment as well as the changes in regulations.
SOP Validation and Continuous Program Improvement
A sedimentation bacteria testing SOP must be validated before routine use and periodically requalified. Validation demonstrates reproducible, accurate results across operators and locations.
SOP Validation Protocol Requirements
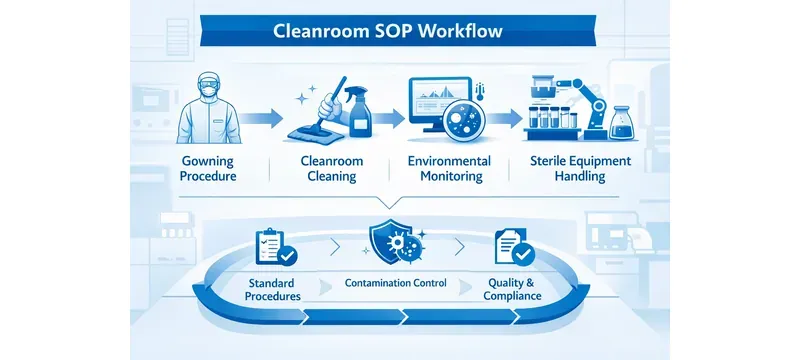
The procedure must comply with the development of the validation master plan specific to the location, as well as complete setting forth the criteria in order to measure acceptability. Among the necessary criteria are coefficient of variation <25% related to reproducibility across operators and the rate of microbial recovery ≥70%.
Reproducibility involves testing done at the same site on the same day using not less than three operators, while recovery verification is achieved with known challenges with microorganisms to arrange colonies on incubation media.
The documentation checklist delivers:
Trend Analysis and Continuous Improvement
In this section of trend review, there will be conducted the analysis on a monthly basis related to the changes within colony counts per specific location and grade, using the moving average control chart. Use moving average control charts to detect gradual shifts in baseline counts that single-point measurements may miss.
Table 4: Continuous Improvement Scorecard
| Metrics | Goals (Targets) | Frequency of Evaluation |
|---|---|---|
| Out-of-Specification (OOS) Rate | Under 2% of all plates | Monthly |
| Risks of Exceeding an Alert Level | Under 5% of all plates | Monthly |
| Trend Analysis | ±20% of deviation from established benchmark | Every 3 months |
| Standard Operating Procedure (SOP) Training | 100% of operating personnel trained | Once a year |
| Validation Frequency | Within requalification time frame | Once a year |
| Metrics | Target | Frequency |
|---|---|---|
| OOS Rate | Under 2% of plates | Monthly |
| Alert Level Exceedance | Under 5% of plates | Monthly |
| Trend Analysis | ±20% deviation | Every 3 months |
| SOP Training | 100% trained | Once a year |
| Validation Frequency | Within requalification frame | Once a year |
Frequently Asked Questions
How often should sedimentation bacteria testing be performed?
Frequency can vary depending on risk level. Grade A requires continuous testing during critical processing. Grade B requires testing on every shift. Grade C typically means 2 to 3 times per week, and Grade D requires weekly testing.
Depending on the risk assessment, frequencies may be adjusted after deviations; however Grade A and B minimum frequencies remain unchanged [6].
What is the difference between settle plate and active air sampling?
Settle plates passively collect bacteria through gravity, reported as colony-forming units (CFU) per plate per exposure time. Active sampling collects known sample volumes of air and reports in CFU per cubic meter.
Class A and B areas require both methods for good manufacturing practice (GMP) environmental monitoring.
What media is required for settle plate method cleanroom testing?
Tryptic soy agar is the standard media utilized for bacterial recovery from settle plates incubated at 30-35°C for 3-5 days. Programs may include additional incubation between 20-25°C for 5-7 days in order to recover fungi and yeasts. Alternative media types must be validated.
How do you calculate CFU per square meter from settle plates?
For a plate with 90 mm, its surface area is 0.00636 square meters. To get CFU per square meter, divide your colony count by its surface area.
Normalize proportionately if exposure time is lesser than four hours as four hours represents deposition of colony within it [2].
What are the GMP limits for viable particle count pharmaceutical settings?
According to EU GMP Annex 1, it has put action limits for Grade A as zero growth, Grade B as five CFU per four hour limit, Grade C as fifty CFU per four hour limit and Grade D as 100 CFU per four hour limit.
Similar principles are used by FDA but the numerical limits are not the same. For instance, alert limits are fixed at 50 and 60 percent of action limits [6].
Can sedimentation testing replace active air sampling entirely?
It is not possible since the regulatory system demands both methodologies to be utilized in grades A and B areas.
Settled plates will help you in getting low cost coverage that will remain consistent throughout the process, whereas, air sampling method supports volumetric count depending on quantitative trends. Both methods need to work together in order derive what is necessary for compliance purposes [7].
References
- [1] iso 14644-1:2015, Cleanrooms and associated controlled environments — Part 1: Classification of air cleanliness by particle concentration.
- [2] ISO 14644-2:2015, Cleanrooms and associated controlled environments — Part 2: Monitoring to provide evidence of cleanroom performance.
- [3] USP <1116>, Microbiological Control and Monitoring of Aseptic Processing Environments.
- [4] USP <797>, Pharmaceutical Compounding — Sterile Preparations.
- [5] EU GMP Annex 1, Manufacture of Sterile Medicinal Products, 2022 revision.
- [6] EU GMP Annex 1 Table 6, Maximum action limits for viable particle contamination, 2022 revision.
- [7] PIC/S GMP Guide Annex 1, Manufacture of Sterile Medicinal Products.
- [8] Napoli C, et al. Air sampling procedures to evaluate microbial contamination: a comparison between active and passive methods. BMC Public Health, 2012.
- [9] FDA 21 CFR Part 210/211, Current Good Manufacturing Practice for Finished Pharmaceuticals.
- [10] GB 50073-2013, Code for design of clean rooms.
- [11] GB 51110-2015, Code for construction and acceptance of cleanroom.