+86 13638632902
 Jason@cleanroomequips.com


 MENU
These standards together determine the classification limits, operational protocols, maintenance intervals, and documentation responsibilities for a facility that conforms with regulatory requirements.
The aim of this article is to provide a systematic reference that takes into account the regulatory requirements for GMP clean and sterile rooms, and the use and management of gmp clean rooms.
It also covers sterile room operation protocols, the maintenance standards for GMP clean rooms, the guidelines of GMP clean room SOPs, and the certification of gmp cleanrooms.
The audience of this article consists of quality managers, clean room engineers, validation specialists, and aseptic processing supervisors in the pharmaceutical sector.
The data that Jason.peng has contributed to the article are based on actual data from Deiiang™, which is a supplier of cleanroom and air filter equipment.
Table 1: Core GMP Clean and Sterile Room Reference Framework
| Standard | Scope | Key Requirement |
|---|---|---|
| EU GMP Annex 1 | Sterile Medicinal Products | Grade A-D classification, CCS, and continuous monitoring |
| ISO 14644-1:2015 | Cleanroom Classification | Limits of particle concentration for ISO classes 1-9 |
| FDA 21 CFR 211 | Drug Manufacturing | Aseptic processing in iso 5 dynamic condition |
| USP 797 | Sterile Compounding | ISO 5 primary engineering control with a recertification every 6 months |
| PIC/S GMP Guide | International Harmonization | Equivalent GMP Standards for member authorities |
| GB 50073-2013 | Chinese National Cleanroom Design and HVAC Code | National code for cleanroom construction and HVAC |
| Standard | Scope | Key Requirement |
|---|---|---|
| EU GMP Annex 1 | Sterile Medicinal Products | Grade A-D classification, CCS, and continuous monitoring |
| ISO 14644-1:2015 | Cleanroom Classification | Limits of particle concentration for ISO classes 1-9 |
| FDA 21 CFR 211 | Drug Manufacturing | Aseptic processing in ISO 5 dynamic condition |
| USP 797 | Sterile Compounding | ISO 5 primary engineering control with a recertification every 6 months |
| PIC/S GMP Guide | International Harmonization | Equivalent GMP Standards for member authorities |
| GB 50073-2013 | Chinese National Cleanroom Design and HVAC Code | National code for cleanroom construction and HVAC |
Core GMP Sterile Room Standards and Regulatory Framework
The global GMP sterile room standards all center around one theme: control of contamination has to be proven and not assumed.
The EU, FDA, WHO, and PIC/S frameworks all have similar ideas in classification, monitoring responsibilities, and documentation requirements.
The EU GMP Annex 1 (2022 version) has the most specific sterile-specific guidance. It requires the contamination Control Strategy to address facility design, HVAC, personnel, and materials.
ISO 14644-1 form the particle concentration basis on which Annex 1 Grade A–D takes place. USP 797 governs sterile compounding in pharmacies requiring ISO 5 for critical areas.
FDA 21 CFR 211 requires the production of sterile drugs in ISO 5 or better conditions.
International GMP Standard Benchmark Comparison
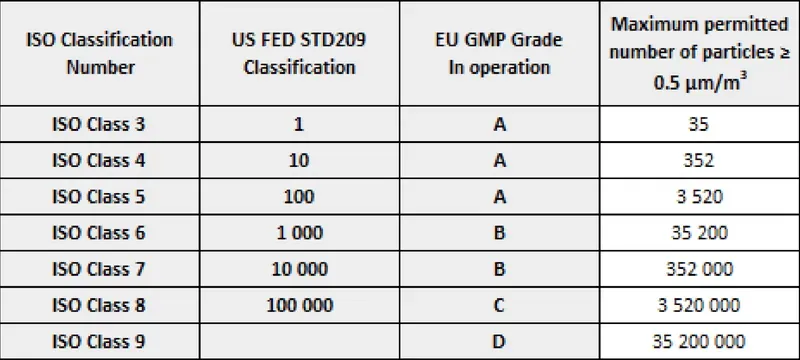
The below table compares the six frameworks classification, monitoring, and requalification. There is consistency in Grade A and ISO 5 conformable applications in the EU, FDA and PIC/S.
WHO GMP creates a risk-based model that is flexible and useful for countries that have less resources.
Table 2: International GMP cleanroom standard benchmarks
| Standard | Classification system | Grade A/critical zone equivalent | Requalification interval | Main monitoring concern |
|---|---|---|---|---|
| EU GMP Annex 1 | Grade A-D | ISO 5: 3,520 particles/m³ ≥0.5 µm | 6 (A/B) and 12 months (C/D) | Continuous viable and non-viable |
| FDA 21 CFR 211 | ISO 5 dynamic | ISO 5 in operation | Periodic (risk-based) | Process validation and sterilization |
| PIC/S GMP Guide | Grade A-D (Annex 1) | ISO 5 | 6 months (A/B) | Same as EU Annex 1 |
| WHO GMP | Grade A-D | ISO 5 | risk-based | CCS and trend analysis |
| ISO 14644-1 | ISO Class 1-9 | iso class 5 | 6 months (ISO 5) and 12 months (ISO 6+) | Particle concentration only |
| USP 797 | ISO 5 (PEC)/7 (buffer area)/8 (ante-area) | ISO 5 (PEC) | 6 months | Viable surface and air sampling |
| GB 50073-2013 | Chinese national code | Class 100 equivalent | As per project specifications | Particles and pressure differential |
| Standard | Grade A equivalent | Requalification interval |
|---|---|---|
| EU GMP Annex 1 | ISO 5: 3,520 particles/m³ ≥0.5 µm | 6 (A/B) and 12 months (C/D) |
| FDA 21 CFR 211 | ISO 5 in operation | Periodic (risk-based) |
| PIC/S GMP Guide | ISO 5 | 6 months (A/B) |
| WHO GMP | ISO 5 | risk-based |
| ISO 14644-1 | ISO Class 5 | 6 months (ISO 5) and 12 months (ISO 6+) |
| USP 797 | ISO 5 (PEC) | 6 months |
| GB 50073-2013 | Class 100 equivalent | As per project specifications |
EU Annex 1 requires a more detailed monitoring process that incurs higher costs for instrumentation but reduces the risk of non-compliance during audits.
FDA 21 CFR 211 has a more flexible risk-based monitoring approach but requires much more detailed validation data. PIC/S member authorities accept Annex 1 alignment as equivalent to local GMP requirements.
EU GMP Annex 1 Cleanroom Requirements Breakdown
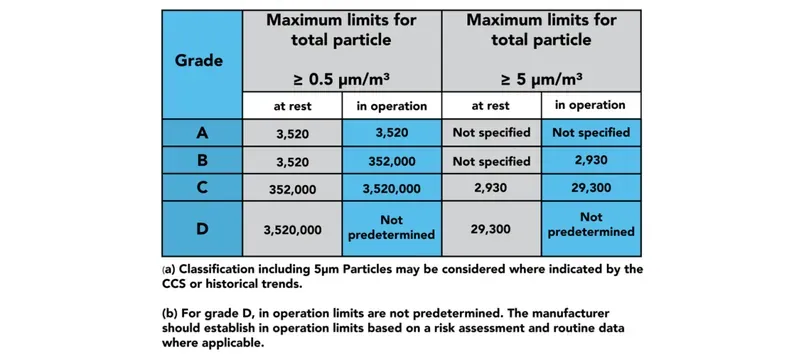
Grade A represents the most critical zone in terms of filling and open product handling operations. Unidirectional laminar airflow is required in the Grade A area with a velocity in the range of 0.36 to 0.54 m/s at the working position.
According to Jason.Peng, the designer of cleanroom products at Deiiang, this velocity is measured at the working position (0.8 to 1 meter above the floor) rather than at the filter.
A common error made in designing the critical zone is having an uneven airflow velocity create eddies and increase the number of particles at active filling points.
The EU GMP Annex 1 cleanroom requirements for Grade A in terms of particle count are ≤ 3,520 particles/m³ at ≥0.5 µm and ≤ 20 particles/m³ at ≥5 µm, both during the operating phase and at rest.
There are also requirements for Grade B, grade c, and Grade D, which need to have particle counts based on ISO 5 and ISO 7 standards.
- Grade A: unidirectional airflow, 0.36–0.54 m/s, 3,520 particles/m³ ≥0.5 µm
- Grade B: ISO 5 at rest, ISO 7 in operation, background to Grade A
- Grade C: ISO 7, less critical aseptic steps
- Grade D: iso 8, closed-system handling and preparation
CCS required for Grade A cleanrooms (housing facilities, HVAC, personnel, materials, and monitoring) must include a formal failure modes and effects analysis-style risk assessment of the sources of particle and microbial contamination mapped to control measures and a tiered monitoring program.
The CCS must be a living document that is updated on a yearly basis. Missing mapping of microbial ingress risks is among the top three findings in EU GMP audits.
GMP Clean Room Use and Management Fundamentals
Operations and management in GMP clean rooms must follow procedures on personnel behavior patterns, access control, and material transfer which are grounded on principles of discipline.
They are generally strict given that any entry into the sterile environment is regarded as a contamination incident. Thus, access is limited to authorized persons only.
The boundaries of operations should be designated through the application of physical barriers as well as enforceable protocols for all transitions.
- Access is granted only to authorized and properly trained personnel.
- Material transfer follows established and recognized decontamination protocols.
- Waste handling uses specially designated and one-way waste pickup channels so that cross-contamination does not occur.
Aseptic Processing cleanroom management Principles
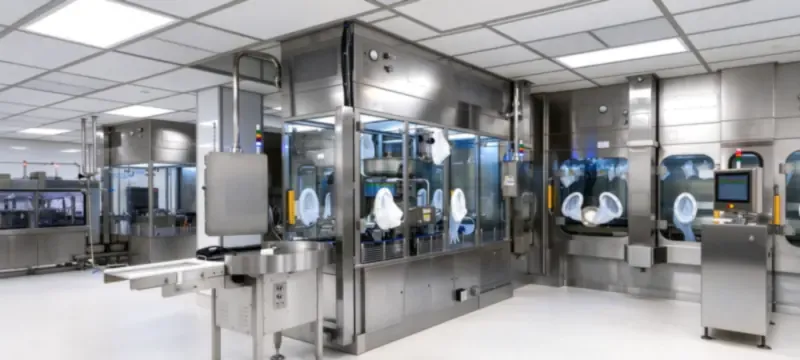
Management of aseptic processing clean rooms must adhere to four basic principles: contamination-first design, unidirectional workflow, pressure cascade compliance, and clear zone signage.
All major decisions in the process are based on the principle of contamination-first, which prioritizes particle and microbial control over production convenience.
- Unidirectional workflow: the principle states that traffic in the clean room is one way only.
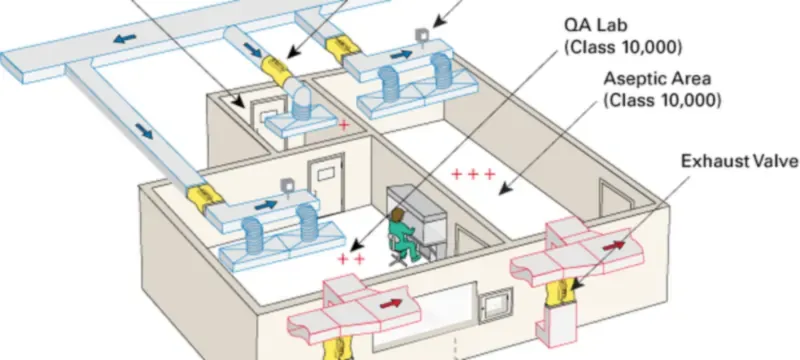
- Pressure cascade compliance: compliance with pressure-cascade rules requires maintenance of a difference of 10-15 Pa between the different operating environments.
- Information signage: indication of real-time grade information, pressure information, and access control status.
Personnel and Material Flow Control Systems
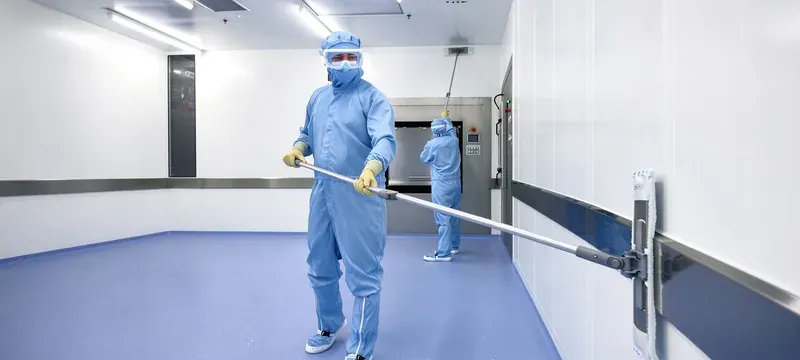
Personnel flow is organized through a multi-level gowning process conducted in clean changing rooms.
- Step 1: Outerwear takeaway and hygiene of hands.
- Step 2: Underclothing and Stage 1 dressing.
- Step 3: Sterile robe, cap and mask put on.
- Step 4: Putting the gloves on and final check visually.
Every 30 minutes gloves need to be changed in Grade A places or as soon as touching anything that is non-sterile. The most frequent gowning failure is adjusting of the mask after final check.
Deiiang on-site service data shows that approximately 40% of viable microbial excursions are due to gowning protocol mistakes as per audit records done by the company.
Material transfer takes place in airlocks that have interlocked doors and only one door cannot be opened at the same time. Decontamination can be wiping, pass-through sterilisation, or UV treatment based on the type of material.
- Airlock rule: interlocked doors, one door open at the same time, a minimum of 30 seconds for the cycle.
- Material decontamination: 70% IPA or sporicidal wipe before transfer.
- Waste removal: sealed containers, separate exit route, and there is no flowback.
Information from Deiiang field service data indicates that interlock door malfunction causes 28% of unplanned pressure cascade excursions.
Jason.peng recommends monthly interlock cycle testing and backup power supplies to avoid unregulated door openings during power fluctuations.
Sterile Room Operation Protocols for Daily Aseptic Activities
Sterile room operation protocols consist of a pre-operating checkup, in-process controls, post-operation shutting down, and batch record documentation.
Each step requires specific acceptance criteria, and any deviation from the acceptance criteria will trigger a documented investigation.
Table 3: Summary of Daily Sterile Room Operational Protocol
| Phase | Main Actions | Acceptance Criteria | Record |
|---|---|---|---|
| Pre-Operation | Counting Particles and verifying pressure | Grade A: ≤3,520/m³ ≥0.5 µm | Monitoring environmental data logger |
| Pre-Operation | Sanitizing equipment | No particles on surfaces, contact with equipment through IPA complete | Sanitization log |
| In-Process | Monitoring viable material in air | Grade A: <1 CFU/m³ | Records of microbial monitoring |
| In-Process | Differential pressure measures | ≥10 Pa between each grade | HVAC records |
| Post-Operation | Cleaning surfaces | All surfaces touched wiped | Records of cleaning |
| Post-Operation | Review the batch record | All problems were noted | Batch record |
| Phase | Main Actions | Acceptance Criteria |
|---|---|---|
| Pre-Operation | Counting Particles and verifying pressure | Grade A: ≤3,520/m³ ≥0.5 µm |
| Pre-Operation | Sanitizing equipment | No particles on surfaces, contact with equipment through IPA complete |
| In-Process | Monitoring viable material in air | Grade A: <1 CFU/m³ |
| In-Process | Differential pressure measures | ≥10 Pa between each grade |
| Post-Operation | Cleaning surfaces | All surfaces touched wiped |
| Post-Operation | Review the batch record | All problems were noted |
Pre-Operation Startup and Verification Protocols
The following checklist can be used directly at the cleanroom entrance with either a pass or failed value to each line item before production occurs.
Table 4: Pre-Operation startup verification checklist
| Checklist Items | Pass/Fail |
|---|---|
| Confirm HVAC has operated for at least 30 minutes before entry | |
| Pressure differentials between adjacent areas have been verified | |
| The calibration of your particle counter has been checked as well as the zero reading | |
| Clean and sanitize all work surfaces using a validated sporicidal agent | |
| The verification of the gowning room supplies and garment integrity | |
| Conduct pre-use HEPA filter visual inspection for damage or leakage | |
| Record all values in the environmental monitoring log | |
| Confirm cleanroom recovery time meets 15–20 minute Annex 1 limit |
| Checklist Items | Pass/Fail |
|---|---|
| Confirm HVAC has operated for at least 30 minutes before entry | |
| Pressure differentials between adjacent areas have been verified | |
| The calibration of your particle counter has been checked as well as the zero reading | |
| Clean and sanitize all work surfaces using a validated sporicidal agent | |
| The verification of the gowning room supplies and garment integrity | |
| Conduct pre-use HEPA filter visual inspection for damage or leakage | |
| Record all values in the environmental monitoring log | |
| Confirm cleanroom recovery time meets 15–20 minute Annex 1 limit |
In-Process Operational Control Requirements
In-process controls require continuous particle monitoring in Grade A and periodic monitoring in Grade B. Viable monitoring uses active air sampling and surface contact plates.
Intervention minimization means every entry into the critical zone must be justified and recorded. Aseptic technique prohibits rapid movements, talking over open product, and reaching across the critical zone.
Any intervention exceeding five seconds requires an immediate impact review by QA, including real-time particle and viable monitoring data captured during the event.
Three or more unplanned interventions per batch trigger a formal CAPA and a process efficiency review. Alert and action limit breaches require the same escalation path.
GMP Clean Room Maintenance Standards and Routine Service
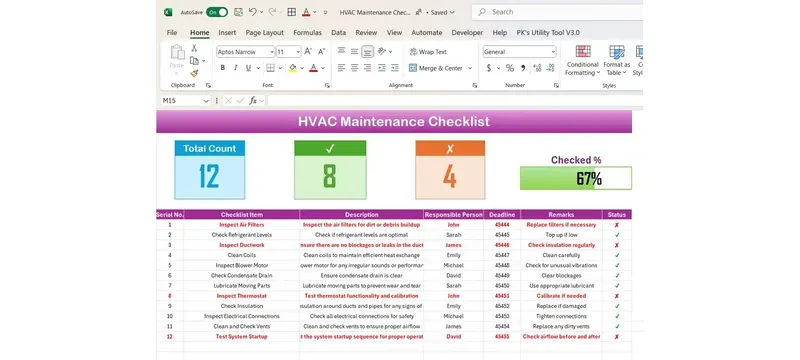
GMP clean room maintenance standards define scheduled service for HVAC, HEPA filters, sanitisation, and instrumentation. Preventive maintenance costs less than remediation.
One failed HEPA filter can invalidate a batch and trigger a regulatory observation.
HEPA filter integrity testing follows iso 14644-3:2019 [6] with DOP (dioctyl phthalate) or PAO (polyalphaolefin) aerosol challenge. HVAC maintenance covers air handling units, ductwork, dampers, and controls.
Sanitisation frequency depends on grade and activity level. Instrument calibration covers particle counters, pressure transducers, and temperature sensors.
Table 5: Responsible Maintenance Schedule of GMP Clean Room
| Item | Activity | Frequency | Acceptance Criterion |
|---|---|---|---|
| HEPA Filter | DOP/PAO SAT | 6 Months (Type A/B), 12 Months (Type C/D) | Penetration = <0.01% |
| HVAC system | air changes Rate Verification | 12 Months | Grade A: 300–600 ACH (laminar) |
| Differential Pressure Monitoring | Continuous | Real Time | Greater than or equal to 10 Pa between grades |
| Sanitization | Daily (Grade A/B), Weekly (Grade C/D) | Daily or Weekly | No viable count above limit allowed |
| Particle Counter Calibration | Calibrated every 12 Months | 12 Months | Within ±10% of reference readings |
| Duct Work | Visual Inspections | 3-6 Months | No dust or damage |
| Pre-Air Filter Replacement | 3-6 Months or ΔP >250 Pa | 3-6 Months or 250 Pa | F7/F8 (EN 779 Legacy / ISO 16890) |
| Item | Frequency | Acceptance Criterion |
|---|---|---|
| HEPA Filter | 6 Months (Type A/B), 12 Months (Type C/D) | Penetration = <0.01% |
| HVAC system | 12 Months | Grade A: 300–600 ACH (laminar) |
| Differential Pressure Monitoring | Real Time | Greater than or equal to 10 Pa between grades |
| Sanitization | Daily or Weekly | No viable count above limit allowed |
| Particle Counter Calibration | 12 Months | Within ±10% of reference readings |
| Duct Work | 3-6 Months | No dust or damage |
| Pre-Air Filter Replacement | 3-6 Months or 250 Pa | F7/F8 (EN 779 Legacy / ISO 16890) |
Filtration and HVAC Preventive Maintenance Schedules
Deiiang H13/H14 HEPA Filters (EN 1822) are constructed from ultrafine glass and elastomer molded frame and polyurethane sealant.
The catalogue provides an efficiency of H14 of 99.995 until 99.999% down to 0.3 micron with an operating temperature of 70° C and 80% humidity maximum.
Example 592 by 592 by 292 mm unit, air delivery is 2500 m³/h with a media area of 20.04 m². It has an initial resistance of ≤220Pa for the H14 cartridge and a final resistance of 400 to 600 Pa. Dust holding capacity is approximately 1200 g.
In a pharmaceutical packaging cleanroom project (class 1000 / ISO 6 target), Deiiang supplied low-profile H14 combined filters, F7 pre-filters, and silicone-free gaskets. Each unit underwent DOP scan testing.
Results from the project showed that particle counts have lessened from an ISO 7 starting point and on to a steady ISO 6, as a consequence of the fact that the low initial total pressure drop led to measurable energy savings.
The DOP-filtered product is able to adequately be used in cleanroom environments where there is a limitation on the constructed height of the room.
Official documentation for the 610*610*150 mm type shows it has a capacity of 2500 m³/h, has a total media area of approximately 22.14 m², and a total dust holding capacity of about 1350 g.
From data provided by the company Deiiang, it was seen that the H14 filter units used in standard Grade B areas display a monthly pressure increase of 0.25-0.35 Pa.
To figure this information, the calculation service life (in months) = (final resistance rating - initial resistance value) / (ΔP value per month) can be used.
Jason.peng recommended that 80% of the rated final resistance of the H14 be utilized in practice since it eliminates the problem of batch contamination from excessive use of this filter.
Environmental Sanitation and Monitoring Maintenance
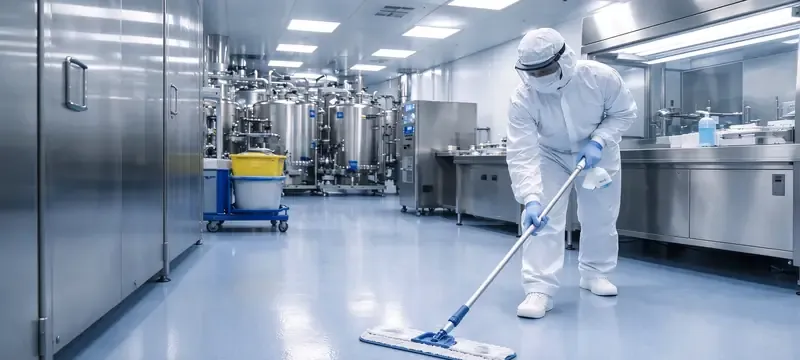
The alternating of sanitizing agents is vital to allow for the avoidance of microbial resistance. A standard rotation schedule alternates sporicidal use on weeks 1 and 3 with non-sporicidal use on weeks 2 and 4, with monthly surface contact plate verification.
Sporicidal agents include 6% hydrogen peroxide and peracetic acid; common non-sporicidal options are 70% IPA and quaternary ammonium compounds.
- Testing of contact plates on the surface: ≥20 locations per Grade A/B zone on a monthly basis.
- Cleaning of the floor: validated mop heads, changed after every 20 square meters per day in Grade A/B.
- Deiiang suggests not using cleaners based on bleach on vinyl or powder-coated wall panels as it leads to discoloration and degradation of surfaces over time.
- Management of waste: use of containers with lids, disposal at least two times during each shift in Grade A.
- Air sampling: active volumetric air sampling, done on a continuous basis in Grade A during operations.
- Trend analysis: review of the trend of viable count every month and in case there is an upward shift, investigate it.
GMP Cleanroom SOP Guidelines for Documented Compliance
GMP cleanroom SOP guidelines provide written guidance that governs every controlled activity taking place in the cleanroom. If SOPs were not there, compliance would be a function of memory.
In the presence of SOPs, compliance depends on documents evidencing actions done and is open to audit.
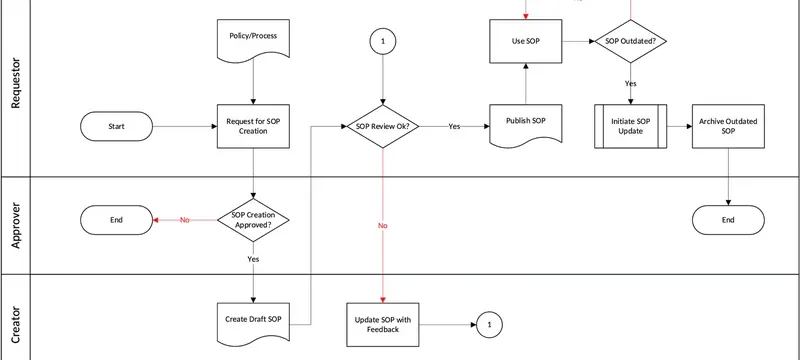
SOPs have a hierarchy that runs from master documents through to specific work instructions. No SOP can be put into use without being signed off by the quality assurance team.
Revision control captures all changes that have been made to the SOPs including reasons and dates.
SOP Document Structure and Approval Workflow
- The header includes the document number, version, effective date, and approval signatures.
- Reason and scope describe the purpose of SOP and where it applies.
- Responsibilities highlight specific roles according to which the SOP must be executed.
- The procedure is written step by step in imperative form.
- Records of retention indicate the forms to be used and the period for which records are to be kept.
- Revision control provides a change history of every change made including the person making the change, the person approving the change, and the reasons for the change.
Each SOP requires a header with document number, version, and effective date. Purpose and scope define applicability. Responsibility assignment names specific roles, not departments.
Record retention rules specify retention period and storage location. Change history logs every revision with author, approver, and rationale.
Validation and Requalification SOP Requirements
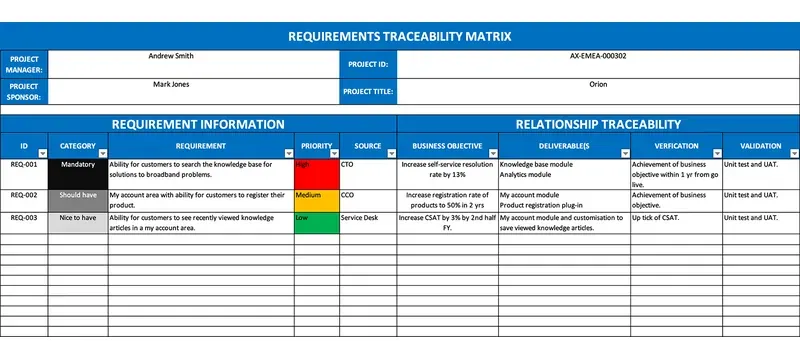
Validation shall be performed in three phases. Installation Qualification (IQ) inspects the equipment to confirm that it conforms to specifications and drawings.
Operational Qualification (OQ) tests the functioning of the system under specified parameters. Performance Qualification (PQ) establishes steady state performance under production conditions.
Requalification shall take place every year or when there has been a change in HVAC, room layout, or process. The HEPA integrity tests shall follow ISO 14644-3, with the aerosol challenge test and a scanning photometer on upstream and downstream.
The airflow pattern test shall be conducted by means of smoke studies to verify the unidirectional airflow in the Grade A.
The requalification frequency for Grade A/B will be six months while that for Grade C/D will be twelve months [7]; the recommendations in ISO 14644-2:2015 [5] set the maximum requalification frequency of six months for ISO 5, and twelve months for ISO 6 and higher.
cleanroom gmp Compliance Certification and Audit Readiness
Cleanroom GMP compliance certification involves third-party verification and on-site inspection, documentation review, and performance testing.
Certification is not a one-time procedure but is an ongoing effort requiring continuous compliance and monitoring.
For audit readiness, all documentation must be current, calibrated equipment must be available and staff must be trained. Any gaps in any one of these areas can lead to a finding of non-compliance.
Table 6: Compliance Myths versus Facts
| Myth | Fact |
|---|---|
| Certification does not need to be renewed | Certification is valid for 12 months after which a full recertification has to be performed every three years |
| HEPA filters do not need to be tested | DOP/PAO testing has to be conducted |
| Documentation can be modified after an audit | SOP must be approved before the procedure commences |
| Grade A monitoring has only to be carried out on a periodic basis | Continuous Grade A monitoring must take place while the unit is functioning |
| Myth | Fact |
|---|---|
| Certification does not need to be renewed | Certification is valid for 12 months after which a full recertification has to be performed every three years |
| HEPA filters do not need to be tested | DOP/PAO testing has to be conducted |
| Documentation can be modified after an audit | SOP must be approved before the procedure commences |
| Grade A monitoring has only to be carried out on a periodic basis | Continuous Grade A monitoring must take place while the unit is functioning |
GMP Certification Pathways and Inspection Criteria
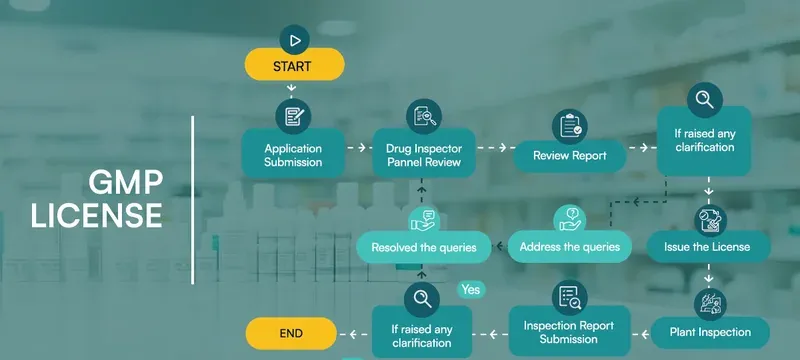
The certification body examines design of the facilities, HVAC efficiency, monitoring and training records.
During inspection, particle count, pressure differentials, and airflow are verified. Review of documentation includes SOP, batch records, and deviation records.
Test for recovery time after generation of particle, verification of filter integrity, verifying air changes takes place.
- Third Party verification: certificate body which has an accreditation in regard to GMP certified cleanrooms.
- On-Site inspection: particle count, pressure, airflow recovery time.
- Documentation review: SOP, validation report, and monitoring records
- Testing for performance: DOP, verification of air changes, recovery test and airflow visualization.
Common Non-Compliance Risks and Corrective Actions
Fill the below self-assessment checklist prior to the inspection or certification visit.
Table 7: Pre-audit compliance self-assessment checklist
| Compliance Element | Self-Check (Yes/No) |
|---|---|
| HEPA integrity test within required interval | |
| SOPs current with signed approval | |
| Pressure differential alarms active and calibrated | |
| All staff training records on file | |
| Open deviation reports closed within 30 days |
| Compliance Element | Self-Check (Yes/No) |
|---|---|
| HEPA integrity test within required interval | |
| SOPs current with signed approval | |
| Pressure differential alarms active and calibrated | |
| All staff training records on file | |
| Open deviation reports closed within 30 days |
According to Deiiang's auditing procedures, HEPA filter maintenance deficiencies are the primary deficiency noted.
HEPA filters that have reached the maximum pressure drop or have exceeded the time frame for integrity testing produce a finding where corrective action must be taken within 7 days.
An example of documentation compliance failures can be described as a lack of signature approvals for SOPs as well as incomplete batch record documentation. A corrective action is electronic document controls and closing all documentation gaps within 30 days.
Training compliance failures exist whenever a new hire is allowed to perform their job functions before going through the required SOP training. Access should be restricted until training is verified, and gaps corrected before re-entry into classified areas.
Frequently Asked Questions
This section answers the frequently asked questions regarding GMP clean and sterile room standards, operations, and compliance related issues.
What are core EU GMP Annex 1 cleanroom requirements for grade A?
Grade A requires constant airflow at 0.36–0.54 m/s and particle limits of 3,520 particles/m³ at ≥0.5 µm and 20 particles/m³ at ≥5 µm. Velocity and uniformity apply instead of a fixed airflow rate. Recovery time after particle generation must not exceed 15–20 minutes.
How frequently do I need to maintain GMP clean room?
Daily surface checks and disinfecting will be required in the cases of Grade A and B. The pressure differential verification on a monthly basis is essential, the semi-annual HEPA testing for the state of Grade A or grade b cleanrooms is essential.
The yearly testing should take place for other grades. The particle counter will be calibrated every year. The inspection of ductworks will be carried out on a visible basis every three to six months.
What should be the constituent elements of GMP cleanroom SOPs?
The SOPs should include purpose, scope, responsibility, and step-wise procedures along with archival rules, and modification procedure. It is necessary to have an approval signature and date in every SOP.
What is the validity period for GMP cleanroom certification?
Certification is valid for twelve months, while the surveillance audit takes place on an annual basis, while a full recertification occurs once every three years. The requalification of both Grade A and B cleanrooms will occur at six-month intervals.
What constitutes best practices for the management of aseptic/sterile processing?
Best practice consists of ensuring all contamination is kept to a minimum level, while as few interruptions as possible occur in the critical area. It is important to have adequate documentation every time the interruption exceeds five seconds.
Are GMP sterile room requirements different in EU and FDA?
There is strong similarity between the principles followed in both organizations. However, the previous EU GMP Annex I document described the use of air velocity rate and other mitigating actions routinely in engaging grade A cleanrooms.
FDA 21 CFR 211 focuses on process validation and sterility assurance, referencing ISO 5 dynamic conditions.
References
- [1] EU GMP Annex 1, Manufacture of Sterile Medicinal Products.
- [2] FDA 21 CFR 211, Current Good Manufacturing Practice for Finished Pharmaceuticals.
- [3] USP General Chapter <797>, Pharmaceutical Compounding—Sterile Compounding.
- [4] ISO 14644-1:2015, Cleanrooms and associated controlled environments—Part 1: Classification of air cleanliness by particle concentration.
- [5] ISO 14644-2:2015, Cleanrooms and associated controlled environments—Part 2: Monitoring to provide evidence of cleanroom performance.
- [6] iso 14644-3:2019, Cleanrooms and associated controlled environments—Part 3: Test methods.
- [7] PIC/S GMP Guide, PE 009-16, Annex 1.
- [8] GB 50073-2013, Code for design of clean room.