+86 13638632902
 Jason@cleanroomequips.com


 MENU
modular cleanroom compliance is achieved through a defined requirements basis, engineered design, controlled construction, commissioning, specified classification and testing, documented operation and monitoring, and qualification only where a regulatory framework requires it. Modular construction does not itself establish ISO classification, GMP compliance where applicable, or a qualified or validated state. It is a delivery method; compliance depends on how requirements are engineered, tested, and maintained.
Understanding Cleanroom Compliance in modular cleanrooms
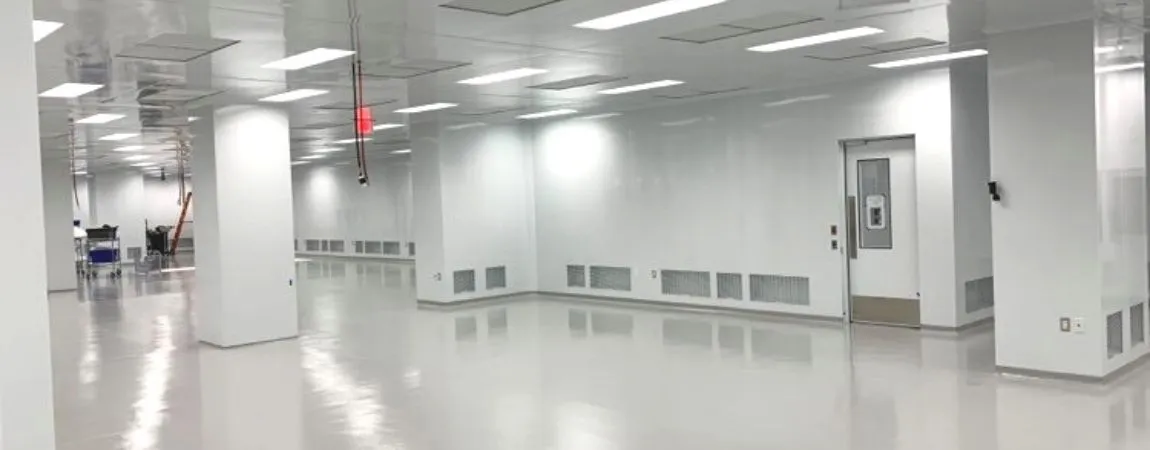
A modular cleanroom enclosure; compliance depends on design, testing, and documentation.
Compliance is a project outcome measured against applicable requirements, not a product label. A modular enclosure can be designed to support a target cleanliness class, but the evidence comes from testing, documentation, and operational control.
It helps to distinguish these related concepts:
| Concept | Purpose | Evidence | Applicable reference |
|---|---|---|---|
| Design conformity | Translate requirements into buildable specifications | Approved drawings, specifications, material schedules | Project requirements/design basis (URS where applicable); iso 14644-4 |
| Commissioning | Verify systems function as specified | Functional test records, balancing reports | Project protocols |
| Particle classification | Determine airborne particle concentration against a class | Particle count report at specified locations and occupancy state | iso 14644-1 |
| Qualification / validation | Demonstrate fitness for intended use within a quality framework | Framework-defined qualification/validation records (e.g., DQ/IQ/OQ/PQ where applicable) | EU GMP Annex 15 — Qualification and Validation (EudraLex Volume 4) where applicable |
| Operational control | Maintain control during routine use | Monitoring data, SOPs, change control records | ISO 14644-2, iso 14644-5 |
Design conformity
Purpose: Translate requirements into buildable specifications
Evidence: Approved drawings, specifications, material schedules
Reference: Project requirements/design basis (URS where applicable); ISO 14644-4
Commissioning
Purpose: Verify systems function as specified
Evidence: Functional test records, balancing reports
Reference: Project protocols
Particle classification
Purpose: Determine airborne particle concentration against a class
Evidence: Particle count report at specified locations and occupancy state
Reference: ISO 14644-1
Qualification / validation
Purpose: Demonstrate fitness for intended use within a quality framework
Evidence: Framework-defined qualification/validation records (e.g., DQ/IQ/OQ/PQ where applicable)
Reference: EU GMP Annex 15 — Qualification and Validation (EudraLex Volume 4) where applicable
Operational control
Purpose: Maintain control during routine use
Evidence: Monitoring data, SOPs, change control records
Reference: ISO 14644-2, ISO 14644-5
ISO 14644-1 classifies air cleanliness by particle concentration. It does not characterize chemical, viable, or other particle properties, and it is not a sterility standard or a GMP certification. Compliance requires the appropriate standard or framework for each purpose.
Which iso 14644 standards Apply—and What Does Each One Actually Cover?
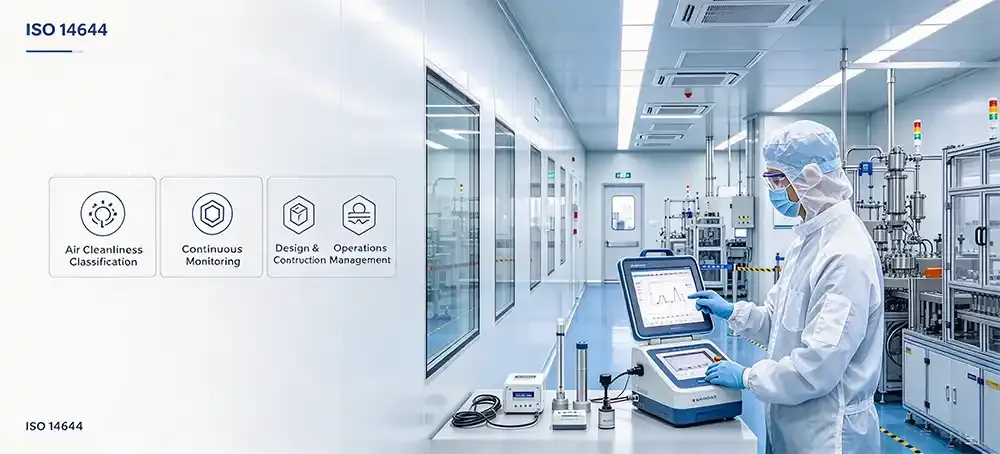
ISO 14644 parts address classification, monitoring, test methods, design, and operations.
The ISO 14644 series addresses different stages. Using the wrong part for the wrong purpose is a common source of confusion.
iso 14644-1:2015 specifies classification of air cleanliness by airborne particle concentration. Only particle populations with threshold particle sizes from 0.1 µm to 5 µm are considered. Light-scattering airborne particle counters measure concentrations at designated sampling locations. Not every class uses the full range.
ISO 14644-2:2015 specifies minimum requirements for a particle-cleanliness monitoring plan. The plan is based on parameters that measure or affect airborne particle concentration.
iso 14644-3:2019 provides test methods for cleanrooms and clean zones. Performance tests are specified for unidirectional and non-unidirectional airflow, and for three possible occupancy states: as-built, at-rest, and operational. The applicable state or states are defined by project requirements. Alternative methods may be used by agreement, but they do not necessarily provide equivalent measurements.
iso 14644-4:2022 specifies the process for creating a cleanroom from requirements through design, construction, and start-up. It applies to new, refurbished, and modified installations.
iso 14644-5:2025 specifies requirements for an operations control programme, covering personnel, entry and exit of personnel and materials, cleaning, maintenance, and monitoring. It does not specifically address biocontamination control; for that topic, iso 14698-1 and iso 14698-2 apply.
How Compliance Is Built Into Modular cleanroom design and Construction
Design review and controlled installation support installed performance.
Compliance begins before fabrication. The project should define the process or use, required ISO air-cleanliness classification where applicable, occupancy state, contamination risks, temperature and humidity requirements, pressure objectives, utilities, applicable codes, and acceptance criteria.
These requirements are translated into the enclosure, airflow and filtration strategy, pressure control, controls system, and utility interfaces. Pressure relationships are project-specific and depend on protection, containment, airflow direction, exhaust arrangements, and adjacent spaces. Positive, negative, or neutral strategies are not universal.
Design reviews should verify interfaces and control changes before construction. Modular construction can shift some fabrication off site, but installation and system integration remain important contributors to installed performance. Maintain approved drawings, material schedules, installation inspection records, sealing records, and change or deviation records.
A concise responsibility matrix helps define who owns each stage:
| Stage | Responsible role (example) | Evidence |
|---|---|---|
| Design approval | Design authority / project engineer | Approved drawings, specifications |
| Installation inspection | Construction quality / site supervisor | Inspection records, sealing records |
| Commissioning | Commissioning agent / controls engineer | Functional test records, balancing reports |
| Classification testing | Test provider / validation specialist | Particle count report, test reports |
| Qualification (where applicable) | Quality unit / validation lead | DQ/IQ/OQ/PQ protocols and reports |
Design approval
Responsible role: Design authority / project engineer
Evidence: Approved drawings, specifications
Installation inspection
Responsible role: Construction quality / site supervisor
Evidence: Inspection records, sealing records
Commissioning
Responsible role: Commissioning agent / controls engineer
Evidence: Functional test records, balancing reports
Classification testing
Responsible role: Test provider / validation specialist
Evidence: Particle count report, test reports
Qualification (where applicable)
Responsible role: Quality unit / validation lead
Evidence: DQ/IQ/OQ/PQ protocols and reports
ISO 14644-4 provides a framework for this process without prescribing specific technological or contractual means.
How Commissioning, Classification Testing and Qualification Should Be Separated
Commissioning, classification, and qualification serve different purposes.
Commissioning is the systematic verification that installed systems function according to agreed specifications. Depending on the agreed scope, it may include checks of HVAC, filtration, pressure control, controls, and utility systems. Commissioning is not classification or qualification.
Particle classification follows ISO 14644-1 and is performed according to the project-specified occupancy state or states. Supporting tests—such as filter integrity, airflow, pressure differentials, recovery time, airflow visualization, and temperature or humidity—are distinct from ISO particle-class designation and are included only when specified. ISO 14644-3 provides test methods for these purposes.
Qualification and validation are framework- and process-specific. They apply only where the quality or regulatory framework requires them. Where DQ/IQ/OQ/PQ stages are used, they are defined by EU GMP Annex 15 — Qualification and Validation (EudraLex Volume 4). These terms are not synonyms for classification or commissioning and should not be imported into industrial cleanroom projects outside such a framework.
An illustrative sequence for many projects runs from design approval, construction, and commissioning, through specified testing and classification, to qualification where applicable, and then release for operation. These stages may overlap.
How Monitoring, Documentation and Change Control Maintain Compliance
Monitoring and documentation support continued operation within specified conditions.
Routine monitoring should be risk-based and scoped to parameters that matter for the room and its use. ISO 14644-2 provides the framework for a particle monitoring plan based on parameters that measure or affect airborne particle concentration. Viable or surface monitoring, if required, comes from another applicable process, microbiological, or regulatory framework—not ISO 14644-2.
ISO 14644-5 provides requirements for an operations control programme, including personnel and gowning, material transfer, cleaning, maintenance, and monitoring procedures. These controls support continued operation within specified conditions.
Documentation, where applicable, may include:
- Approved specifications and as-built drawings
- Commissioning records
- Specified supporting-test reports
- ISO classification report
- Calibration records where applicable
- Standard operating procedures
- Maintenance and monitoring logs
- Change-control records
- Qualification records where applicable
Each record should trace to a requirement, an acceptance criterion, a responsible party, and the current configuration.
A compact six-step compliance-evidence checklist:
| Step | Owner | Evidence | Acceptance point |
|---|---|---|---|
| 1. Requirement/design basis | Project owner | URS or requirements document | Approved requirements |
| 2. Applicable standard/framework | Quality/regulatory | Scope decision record | Framework confirmed |
| 3. Approved design | Design authority | Issued drawings, specifications | Design approval |
| 4. Controlled installation/change | Construction/site QA | Inspection records, change logs | Installation verified |
| 5. Commissioning/testing | Commissioning/test provider | Functional tests, classification report | Tests passed |
| 6. Operations/monitoring | Operations/quality | SOPs, monitoring data | Continued control |
1. Requirement/design basis
Owner: Project owner
Evidence: URS or requirements document
Acceptance: Approved requirements
2. Applicable standard/framework
Owner: Quality/regulatory
Evidence: Scope decision record
Acceptance: Framework confirmed
3. Approved design
Owner: Design authority
Evidence: Issued drawings, specifications
Acceptance: Design approval
4. Controlled installation/change
Owner: Construction/site QA
Evidence: Inspection records, change logs
Acceptance: Installation verified
5. Commissioning/testing
Owner: Commissioning/test provider
Evidence: Functional tests, classification report
Acceptance: Tests passed
6. Operations/monitoring
Owner: Operations/quality
Evidence: SOPs, monitoring data
Acceptance: Continued control
Modifications should trigger an impact assessment. Further tests, reclassification, or qualification depend on the impact of the change.
When Do GMP and Pharmaceutical Requirements Apply?
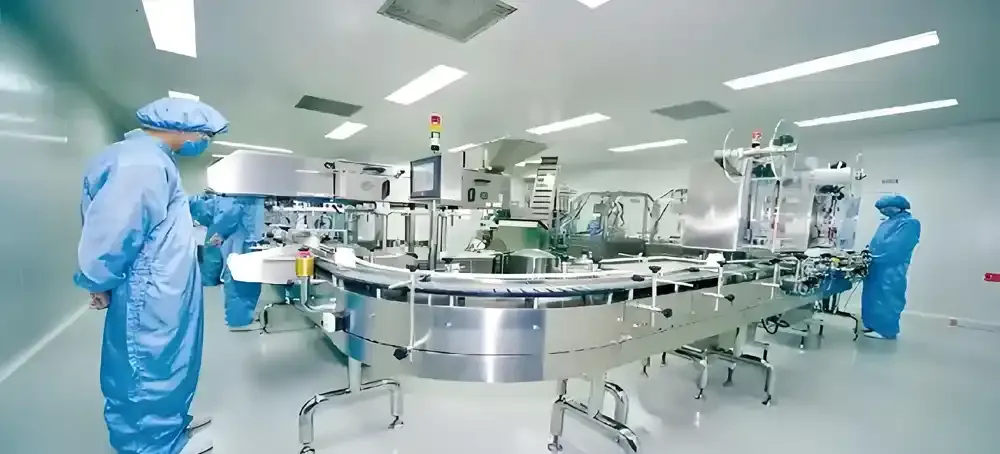
GMP requirements apply where the product and process fall under the relevant medicinal or drug framework.
GMP requirements apply where the product and process fall under the relevant medicinal or drug regulatory framework. They do not apply to every modular cleanroom.
For sterile drug and biological products produced by aseptic processing, FDA guidance issued in October 2004 describes CGMP expectations. EU GMP Annex 1, revised in 2022, specifies requirements for the manufacture of sterile medicinal products. Annex 1 introduces a Contamination Control Strategy that should define critical control points and assess the effectiveness of design, procedural, technical, and organisational controls.
EU gmp grades and ISO 14644 particle classes are different frameworks, not synonyms. ISO classification alone does not establish pharmaceutical compliance. Additional microbiological, personnel, and aseptic expectations apply within applicable sterile-pharmaceutical frameworks.
FDA guidance states that aseptic manufacturers should not rely solely on ISO 14644-1 or ISO 14644-2 for facility qualification.
Deiiang product pages identify offered systems and components only. They are company-published product information, not evidence of compliance or performance.
Frequently Asked Questions
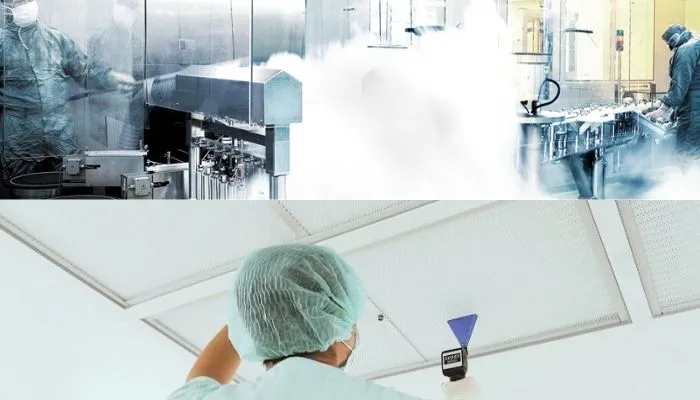
Compliance depends on project requirements and applicable standards or frameworks.
Does a modular cleanroom automatically comply with ISO 14644?
No. Modular construction is a delivery method. Compliance depends on applicable project requirements and the relevant ISO provisions being addressed through design, installation, testing, and documentation.
Is classification the same as qualification or validation?
No. Classification determines airborne particle concentration against a class. Qualification and validation are framework-specific processes demonstrating fitness for intended use where a regulatory framework requires them.
What tests and documents may be needed?
They depend on the project. They may include classification testing at specified occupancy state(s), commissioning records, calibration records, approved drawings, specified supporting-test reports, and qualification records where applicable.
Does every cleanroom need continuous monitoring?
No. ISO 14644-2 provides the framework for a particle monitoring plan based on parameters that measure or affect airborne particle concentration. Monitoring frequency and scope are risk-based and project-specific.
Does GMP apply to every modular cleanroom?
No. GMP applies where the product and process fall under the relevant medicinal or drug regulatory framework. Industrial sites may follow process, customer, ISO, or local requirements rather than a pharmaceutical GMP framework.
Conclusion — Compliance Comes From the Complete System and Its Evidence
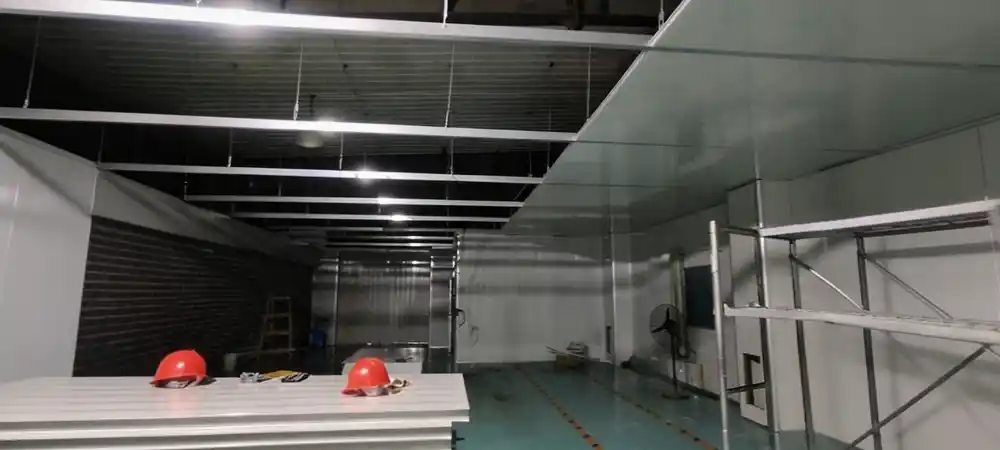
A cleanroom installation illustrates project context; the project evidence determines whether requirements are met.
Modular cleanroom compliance is built through a complete chain: defined requirements, engineered design, controlled construction, commissioning, specified testing and classification, qualification where applicable, and ongoing operational control with documentation. No single product or construction method guarantees compliance. Each project must establish its requirement basis, demonstrate performance through appropriate evidence traceable to requirements and acceptance criteria, and maintain control through monitoring, procedures, and change management.
References
ISO/Technical Standards
- ISO 14644-1:2015 — Classification of air cleanliness by particle concentration
- ISO 14644-2:2015 — Monitoring to provide evidence of cleanroom performance related to air cleanliness by particle concentration
- iso 14644-3:2019 — Test methods
- ISO 14644-4:2022 — Design, construction and start-up
- ISO 14644-5:2025 — Operations
Regulatory/GMP Sources
- FDA Guidance — Sterile Drug Products Produced by Aseptic Processing—Current Good Manufacturing Practice (October 2004)
- EU GMP Annex 1 (2022 revision) — Manufacture of Sterile Medicinal Products
- EU GMP Annex 15 — Qualification and Validation (EudraLex Volume 4)