+86 13638632902
 Jason@cleanroomequips.com


 MENU
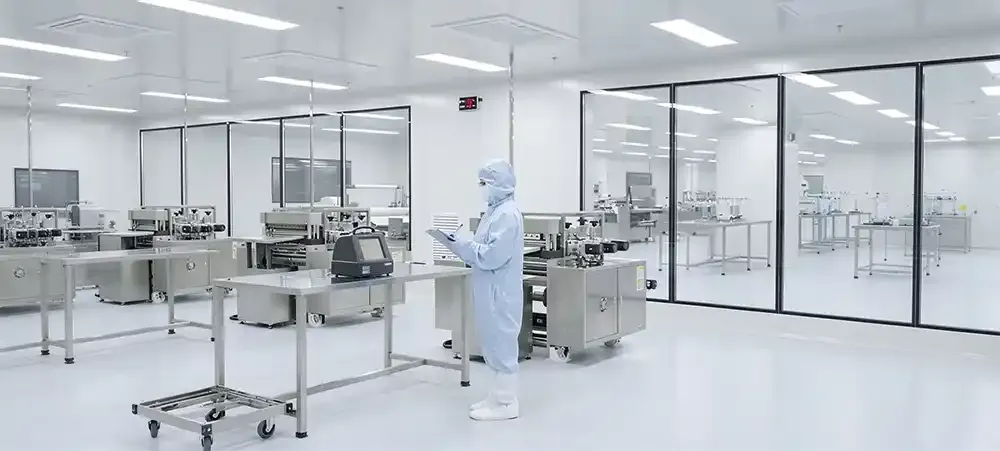
modular cleanrooms made for cosmetology production establishments control pollution caused by employees, equipment, and surroundings. They consist of ready-made housing materials, air-conditioning systems, and filters and they create conditions for production regulating the process. Nevertheless, the mere classification of air contamination level according to ISO standards is not enough for compliance with GMP and there is no universal classification for all production processes in this area.
It is necessary to apply a risk-based approACH to organize the process of cleanroom construction, as it should connect product requirements with pollution sources, cleaning booking methods, and building productivity. The given materials will help learn about the necessary requirements and information about the design of modular cleanrooms including the results of tests and project documentation.
Evidence used in this article. Primary standards and regulations (iso 14644-1, ISO 22716, EU Regulation 1223/2009, U.S. MoCRA) are cited directly in the References. Manufacturer component data comes from supplied Deiiang product sheets and is identified as such. The adjacent-industry reference is a company-reported case page. Everything else is project-specific engineering judgment that must be confirmed for the actual facility.

Why Cosmetics Facilities Need a Risk-Based cleanroom Strategy
The cosmetics industry covers an enormous range of products: water-based emulsions, anhydrous balms, pressed powders, loose powders, gels, and more. A single facility may produce several categories on adjacent lines. Applying one environmental specification to all of them is neither practical nor scientifically justified. The starting point is understanding where contamination actually matters for each product and process.
Identify the product and process risks first
Water-based formulas bring different microbial control challenges than anhydrous products. Anhydrous products are not automatically risk-free, however. Water activity, preservative system, open exposure time, raw material condition, and the way the finished product is used by the consumer all influence the risk profile. Powder products generate dust and require different containment strategies than liquid filling. Hot processing creates vapor and condensation concerns; cold processing may involve longer open hold times. Open transfers of bulk product, filling nozzles, and packaging feed systems each expose product to the environment in different ways. People movements, equipment cleaning, and maintenance activity all introduce variables.
None of this implies that every cosmetics process requires sterile conditions or pharmaceutical-grade environments. Many cosmetics operations function effectively with controlled but non-classified spaces, supported by good hygiene practices and appropriate cleaning regimes. The risk assessment determines the level of control, not an assumption that “cleaner is always better.”
The user requirement specification (URS) should capture the inputs that drive these decisions. A practical starting list includes:
- Formulation water content and water activity
- Open exposure time during each process step
- Batch size and transfer volumes
- Number of personnel and their movement patterns
- Dust, vapor, or volatile emissions from the process
- Cleaning methods and agents used on equipment and surfaces
- Shared equipment between product types
- Cleanliness state of incoming primary packaging
- Finished-product microbiological specification
Define the intended state of control
Before selecting a classification or designing a room, define what “controlled” means for the specific project. This includes the products and process steps in scope, which operations are open versus closed, how people and materials move through the space, the environmental setpoints (temperature, humidity, pressure relationships), cleaning and maintenance procedures, the target market and its regulatory basis, and the acceptance criteria that will be used to verify performance.
Local engineering inputs also shape the design. These include the project country and city, outdoor temperature and humidity design conditions, altitude, voltage and frequency, available chilled and heating sources, building floor-to-floor height, slab and ceiling load capacity, drainage provisions, fire protection strategy, local building and electrical codes, and transport or installation constraints. The values must come from the actual site and project documents, not from generic assumptions.
This definition becomes the basis for design decisions and the benchmark for validation. Without it, the project risks over-specifying some areas and under-protecting others.
Why a universal iso class is not the answer
There is no single ISO class that applies to all cosmetics manufacturing. Controls depend on the product, the process, the risk assessment, and the requirements of the market where the product will be sold. Product characteristics such as water content, preservative system, or alcohol level are risk factors, but they do not by themselves determine an ISO class. The open versus closed nature of the operations, the packaging cleanliness, the personnel flow, and the cleaning regime all contribute.
ISO 14644-1 provides a classification framework, but it does not prescribe which class a given cosmetics process must achieve. That determination comes from the risk assessment and applicable market requirements, not from the standard itself. The next section explains the boundary between classification and GMP more fully.
iso 14644 Classification and Cosmetic GMP Are Different Layers
A common misconception in cosmetics facility planning is that achieving a certain ISO class equals GMP compliance. These are separate concepts that serve different purposes. Understanding the distinction prevents wasted investment and misdirected validation effort. Later sections of this article cross-reference this one rather than repeating the same explanation.
What ISO 14644-1 establishes
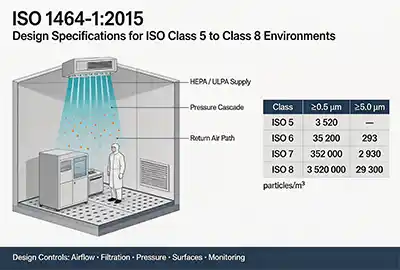
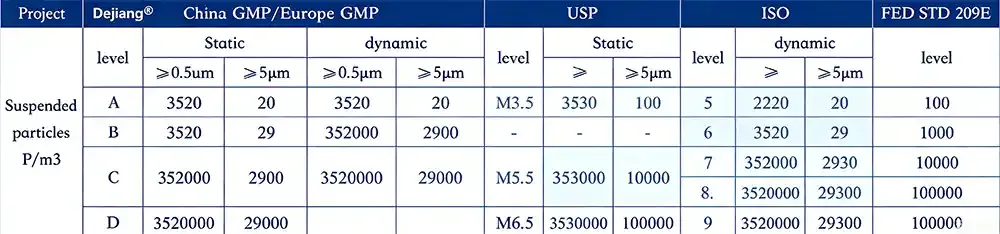
iso 14644-1:2015 specifies the classification of air cleanliness by particle concentration. It uses cumulative distributions based on threshold particle sizes and defines the maximum allowable concentrations for each class. The standard describes classification; it does not prescribe which class a given process must meet, and it does not characterize the physical, chemical, radiological, or viable nature of the particles counted.
In practical terms, ISO 14644-1 tells you how many particles of a given size or larger are permitted in a defined volume of air, under specified test conditions. That number describes air cleanliness. It does not describe whether the space is suitable for a particular cosmetics process. Refer to the published standard for the full classification table and testing provisions.
What ISO 22716 addresses
ISO 22716:2007 provides guidelines for the production, control, storage, and shipment of cosmetic products. It addresses the systems and practices that support consistent product quality, including documented controls across the manufacturing operation. It is broader than particle classification.
Two scope limits matter. ISO 22716 does not cover personnel safety or environmental protection, and it is not intended for research and development or for the distribution of finished products. Adopting the standard does not by itself cover all safety, environmental, and supply-chain obligations a manufacturer may have.
How market obligations change the compliance plan
Regulatory requirements vary by market. The following is an EU-specific example. Article 8 of Regulation (EC) No 1223/2009 requires that the manufacture of cosmetic products comply with good manufacturing practice. Separately, compliance with the relevant harmonized standards can support a presumption of conformity, and EN ISO 22716:2007 is the harmonized standard referenced for cosmetics GMP in the EU context. These are two distinct points: the Article 8 GMP obligation, and the separate role of harmonized standards in supporting a presumption of conformity. This does not mean every EU cosmetics facility must achieve a specific ISO 14644 class; it means the manufacturing operation must follow GMP, and ISO 22716 provides a recognized framework.
In the United States, the Modernization of Cosmetics Regulation Act of 2022 (MoCRA) addresses several areas. It requires facility registration and product listing, and it establishes adverse event reporting obligations. Separately, MoCRA directs FDA to establish GMP requirements for cosmetics facilities. Because FDA rulemaking status changes over time, readers should verify the current status directly with FDA and the Federal Register.
A practical two-layer specification
The following comparison sets out the two layers. It is a comparison framework, not a statement that both layers are required in every project.
| Layer | What it addresses | Typical evidence |
|---|---|---|
| Airborne-particle classification (ISO 14644-1, where applicable) | Air cleanliness by particle concentration at specified sizes, under defined test states | Particle count test reports at agreed locations and conditions |
| GMP controls (ISO 22716 framework) | Premises, hygiene, personnel, materials, production, documented controls, cleaning, training, quality oversight | Procedures, records, training logs, documented cleaning procedures and records, cleaning verification or validation where justified by risk and applicable requirements, audit reports, deviation handling |
- Layer
- Airborne-particle classification (ISO 14644-1, where applicable)
- What it addresses
- Air cleanliness by particle concentration at specified sizes, under defined test states
- Typical evidence
- Particle count test reports at agreed locations and conditions
- Layer
- GMP controls (ISO 22716 framework)
- What it addresses
- Premises, hygiene, personnel, materials, production, documented controls, cleaning, training, quality oversight
- Typical evidence
- Procedures, records, training logs, documented cleaning procedures and records, cleaning verification or validation where justified by risk and applicable requirements, audit reports, deviation handling
Both layers matter, but they answer different questions. Classification answers “how clean is the air?” GMP answers “is the operation controlled and documented to consistently produce quality product?”
Terminology: classification, commissioning, qualification, certification
Classification is the particle-based determination of air cleanliness. Commissioning confirms that installed systems are installed correctly, controlled correctly, and ready for operation. Qualification and validation produce documented evidence under the project quality plan that agreed requirements are met. Certification always depends on the issuing party and the scheme; a certification mark without a named issuer and scheme should not be treated as evidence of compliance.
Modular cleanroom design Around the Cosmetics Process
Once the risk assessment and regulatory basis are clear, design can translate those requirements into physical and mechanical choices. Modular construction offers flexibility for layout changes and expansion, but the design still needs to follow the process, not the other way around.
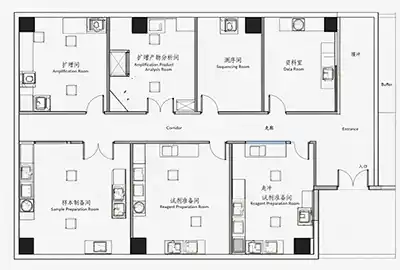
Zoning, layout, and movement
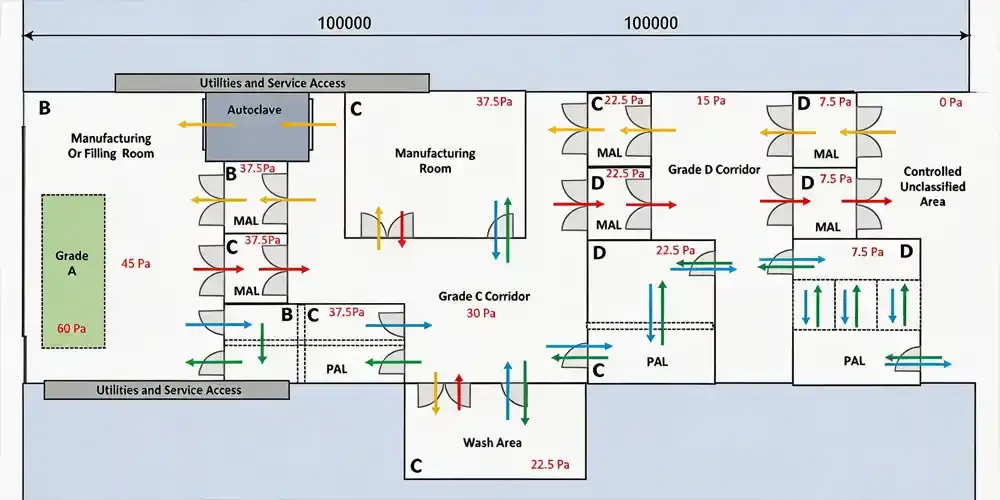
Start by mapping the actual flow of materials and people: receipt of raw materials, weighing and dispensing, compounding or mixing, holding, filling, packaging, and staff entry and exit. Identify where open product is exposed and where cross-contamination risks exist between different product types or between dirty and clean activities.
The goal is to reduce unnecessary crossover between areas of different contamination risk. This does not mean every facility must follow one universal flow pattern. The appropriate zoning depends on the products, the process, and the building constraints. A small facility producing one product category may have a simpler flow than a multi-product site with shared equipment.
A conceptual process flow can be summarized as: material receipt; weighing and dispensing; compounding or mixing; holding; filling; packaging; staff entry and exit with material airlock positions. This is an illustrative conceptual example, not a prescription for any specific project.
The cleanroom HVAC system
HVAC is the primary mechanical system for controlling airborne particles, temperature, humidity, and pressure relationships. Positive pressure is not the default for every area. Powder handling, volatile emissions, odors, and occupational exposure may call for exhaust or reverse pressure differentials. The direction of pressure relationships should follow the product protection and contaminant containment goals defined in the risk assessment.
Key design variables include room volume, occupancy, equipment heat loads, process exhaust, door opening frequency, filter terminal resistance, outdoor design conditions, temperature and humidity loads, and expansion headroom. air change rates, pressure differentials, and filtration levels must be calculated for the specific project. Generic values should not be presented as guaranteed design targets. Final values require engineering calculation and agreement on acceptance criteria during the design phase.
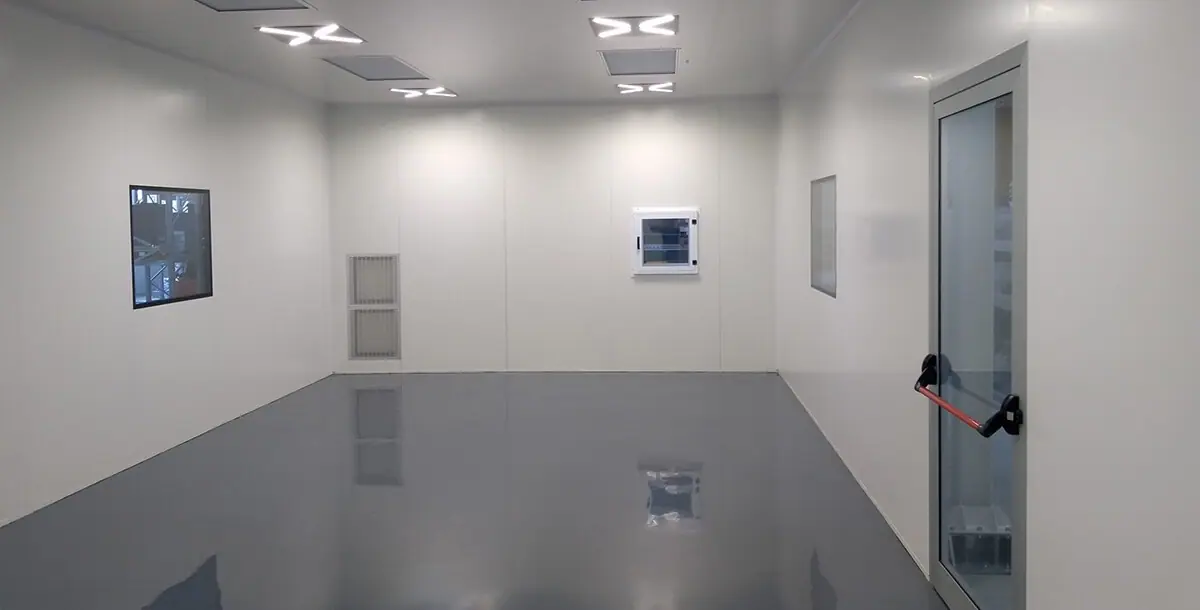
Modular envelope and cleanability
The cleanroom envelope — panels, ceilings, doors, glazing, and joints — should support cleanability, durability, and access. Surfaces need to withstand the cleaning agents and methods used in the facility. Joints and corners should minimize crevices where particles or residues can accumulate. Penetrations for utilities, piping, and equipment should be sealed or otherwise managed. Sealants, windows, and floor systems should be checked for compatibility with the cleaning regime. Antibacterial or corrosion-resistance claims should not be repeated without a supporting test report from the supplier.
Deiiang offers modular cleanroom components and clean booths that address these envelope considerations. The selection between a full-room approach and a localized clean booth should follow from the process risk assessment, not from a default preference for one option. Product information and claims for these systems are available on the manufacturer’s pages.
Full-room control versus localized clean booths
A full modular room provides environmental control across an entire production area. A clean booth or localized enclosure provides control around a specific operation or piece of equipment. The choice is a process-risk decision. The comparison below is a decision aid, not a cost or schedule comparison.
| Consideration | Full modular room | Localized clean booth |
|---|---|---|
| Control scope | Entire production area | Specific operation or equipment |
| Dependence on existing workshop | Lower; envelope provides separation | Higher; performance depends on surrounding environment |
| Personnel flow | Managed across the room | Managed at the booth interface |
| Process interfaces | Multiple connected operations | Single operation or equipment interface |
| Expansion | Requires utility and HVAC headroom | Often easier to relocate or add |
| Verification boundary | Defined by the room envelope | Defined by the booth envelope and surrounding conditions |
| Typical scenario | Multiple sensitive steps or extensive movement | Isolated sensitive step |
- Consideration
- Control scope
- Full modular room
- Entire production area
- Localized clean booth
- Specific operation or equipment
- Consideration
- Dependence on existing workshop
- Full modular room
- Lower; envelope provides separation
- Localized clean booth
- Higher; performance depends on surrounding environment
- Consideration
- Personnel flow
- Full modular room
- Managed across the room
- Localized clean booth
- Managed at the booth interface
- Consideration
- Process interfaces
- Full modular room
- Multiple connected operations
- Localized clean booth
- Single operation or equipment interface
- Consideration
- Expansion
- Full modular room
- Requires utility and HVAC headroom
- Localized clean booth
- Often easier to relocate or add
- Consideration
- Verification boundary
- Full modular room
- Defined by the room envelope
- Localized clean booth
- Defined by the booth envelope and surrounding conditions
- Consideration
- Typical scenario
- Full modular room
- Multiple sensitive steps or extensive movement
- Localized clean booth
- Isolated sensitive step
This is not simply a cost comparison. A booth that is undersized for the operation, or that is placed in an environment that undermines its performance, will not deliver the intended control. The decision should be made with the process and quality teams, based on where control is actually needed.
From Concept to Handover: The One-Stop Project Workflow
A one-stop approach coordinates the activities that would otherwise be split across multiple vendors and disciplines. Scope varies by project. Responsibilities should be confirmed in writing, because a one-stop provider may not cover every activity below. The responsibility table after the workflow steps shows typical responsibility allocation; it is a planning aid, not a claim about any specific provider’s committed scope.
User requirements and concept design
The project starts with a user requirement specification: what the facility needs to do, what products and processes are in scope, what regulatory basis applies, and what constraints exist. A site survey establishes the physical conditions. A process map identifies flows and interfaces. Preliminary zoning defines the controlled areas and their relationships. A utility review identifies power, water, drainage, and other services. A standards register documents the applicable requirements.
A URS input table is a useful working tool. A sample structure may include:
- Product family and formulation characteristics
- Process steps and open/closed status
- Batch size, throughput, and shift pattern
- Environmental targets and acceptance criteria
- Site utilities and building constraints
- Target market and regulatory basis
- Documentation and training requirements
- Expansion and change-control expectations
Detailed engineering and design review
Concept design progresses to coordinated drawings across architectural, mechanical, electrical, and process disciplines. HVAC calculations establish airflows, pressures, and equipment sizing. Equipment interfaces are defined for both utilities and control systems. Maintenance clearances are confirmed. General design review checkpoints include equipment entry routes, maintenance access space, filter replacement access, drainage and threshold conflicts, door-opening pressure disturbances, and process exhaust make-up air. These are general checkpoints, not a record of any specific project.
Manufacturing, preassembly, and quality checks
Modular components are manufactured and, where practical, preassembled before shipment. Inspection and traceability records document that components meet specifications. It is important to recognize that component inspection does not prove finished-room performance. A panel that meets its specification, a filter that passes its test, and an HVAC unit that runs correctly will still require integrated testing after installation.
Installation and commissioning
Installation covers the envelope, mechanical and electrical integration, controls, and interconnection of systems. Commissioning includes balancing of airflows, verification of control sequences, and initial checks of system operation. Snagging identifies and resolves issues before formal testing. Readiness checks confirm that the systems are prepared for qualification activities.
Qualification, testing, and handover
Installation is not the same as verification. Qualification and testing demonstrate that the installed facility performs according to the agreed criteria. The documentation package for handover typically includes approved drawings, operation and maintenance manuals, test reports, calibration records, training records, and maintenance schedules. The scope of testing depends on the project requirements and the regulatory basis.
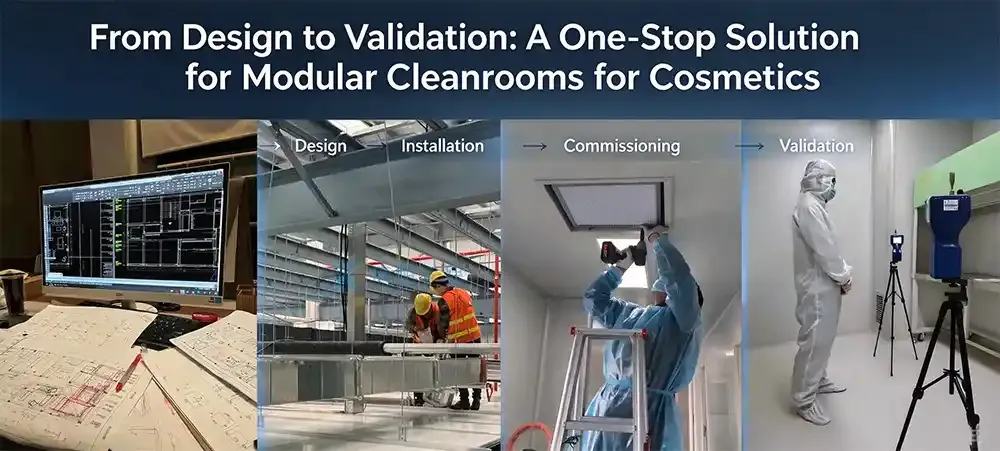
The seven-step workflow below can be used as a planning checklist. It is a conceptual example, not a fixed sequence for every project.
- User requirements and standards register
- Concept design and preliminary zoning
- Detailed engineering and design review
- Manufacturing and preassembly with quality checks
- Installation and commissioning
- Qualification and testing against agreed criteria
- Operational handover with documentation and training
| Activity | Owner / manufacturer | Provider / contractor | Third-party tester | Regulatory advisor |
|---|---|---|---|---|
| URS and acceptance criteria | Lead | Support | — | Support |
| Concept and detailed design | Approve | Lead | — | Review |
| Manufacturing and component inspection | Approve | Lead | Witness where agreed | — |
| Installation and commissioning | Approve | Lead | — | — |
| Qualification and testing | Approve | Support | Lead where engaged | Review |
| Handover documents and training | Approve | Lead | — | — |
- Activity
- URS and acceptance criteria
- Owner / manufacturer
- Lead
- Provider / contractor
- Support
- Third-party tester
- —
- Regulatory advisor
- Support
- Activity
- Concept and detailed design
- Owner / manufacturer
- Approve
- Provider / contractor
- Lead
- Third-party tester
- —
- Regulatory advisor
- Review
- Activity
- Manufacturing and component inspection
- Owner / manufacturer
- Approve
- Provider / contractor
- Lead
- Third-party tester
- Witness where agreed
- Regulatory advisor
- —
- Activity
- Installation and commissioning
- Owner / manufacturer
- Approve
- Provider / contractor
- Lead
- Third-party tester
- —
- Regulatory advisor
- —
- Activity
- Qualification and testing
- Owner / manufacturer
- Approve
- Provider / contractor
- Support
- Third-party tester
- Lead where engaged
- Regulatory advisor
- Review
- Activity
- Handover documents and training
- Owner / manufacturer
- Approve
- Provider / contractor
- Lead
- Third-party tester
- —
- Regulatory advisor
- —
Cost and schedule factors vary by project and should be confirmed against project-specific information rather than assumed. Typical factors include project area and room count, environmental goals, climate, materials, equipment interfaces, site constraints, validation scope, transport, and local construction conditions.
Cleanroom Validation: Proving the Agreed Requirements
Validation demonstrates that the facility performs as intended and meets the agreed acceptance criteria. In this article, cleanroom validation refers to the written activities and evidence agreed for the project. It is not automatically equivalent to product or process validation, and it is not a certification issued by a regulator.
Define acceptance criteria before testing
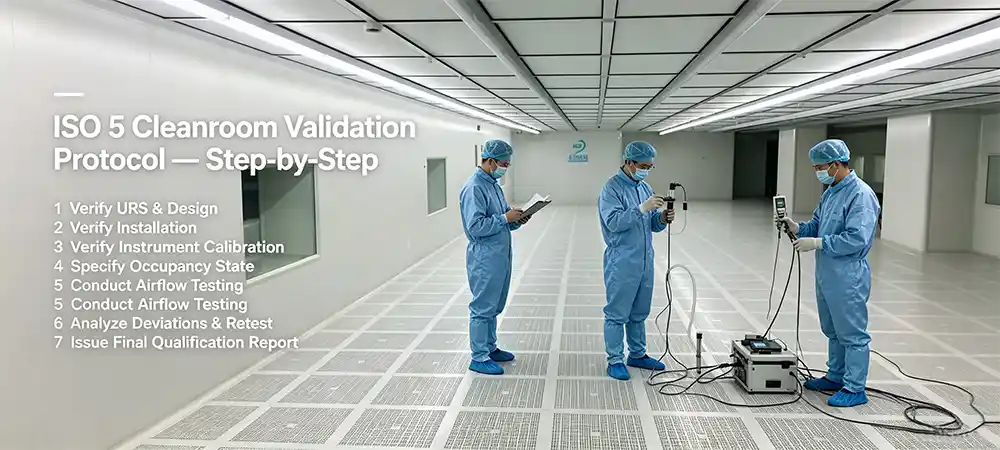
Methods, conditions, sampling locations, instruments, responsibilities, and pass/fail criteria should be agreed before testing begins. This prevents disputes later and ensures that the testing actually addresses the requirements. For particle classification, this includes the test state (as-built, at-rest, or operational) and the particle sizes to be measured.
Typical commissioning and qualification checks
The list below reflects items typically considered. The exact protocol is project-specific, and not every item is mandatory for every cosmetics project.
- Airflow and balancing
- Pressure differential between adjacent areas
- Filter integrity testing where specified
- Temperature and humidity verification
- Airborne particle counting where a classification is specified
- Recovery testing or airflow visualization if required
- Alarm and control function verification
- Documentation review
Why validation is more than a particle-count certificate
A passing particle count confirms that air cleanliness met a defined limit at a specific time and condition. It does not confirm that the facility is GMP-ready. GMP readiness also depends on procedures, cleaning effectiveness, personnel practices, maintenance programs, monitoring plans, training, and change control. A facility can meet a particle classification and still have gaps in documentation, cleaning procedures, or personnel flow.
Maintaining the validated state
Validation is not a one-time event. Preventive maintenance, filter replacement schedules, instrument calibration, trend monitoring, periodic re-testing, deviation handling, and requalification after significant changes all contribute to maintaining the validated state. Without these ongoing controls, a facility may no longer meet its approved performance criteria or quality-system requirements over time.
Adjacent-Industry Reference: What Can and Cannot Be Learned
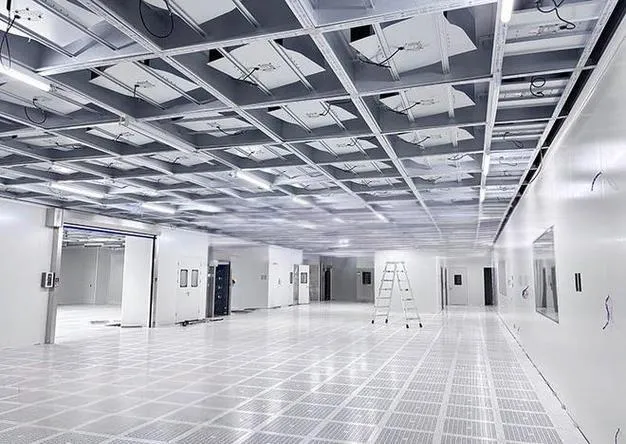
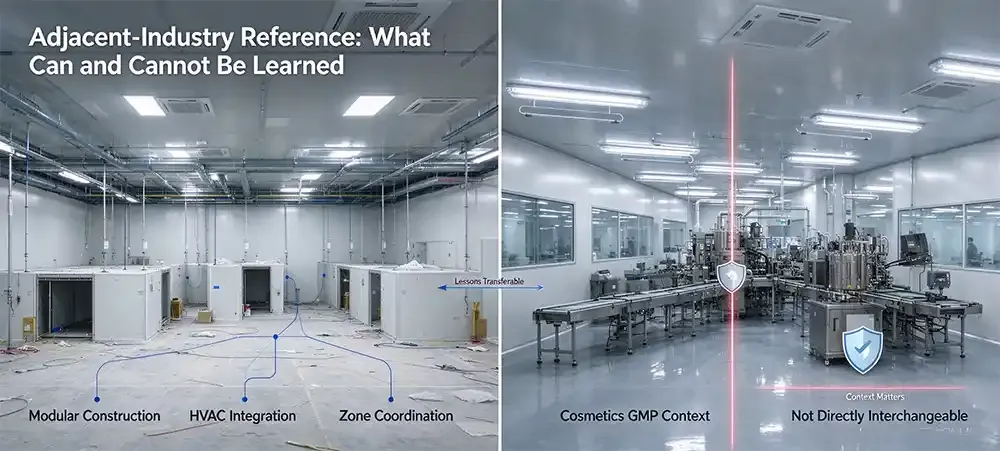
Deiiang’s public case materials include a reported food and solid-beverage facility project. The page describes approximately 8,000 m² and uses the term “class 100,000”. This is presented as a Deiiang-reported food/solid-beverage reference. It is not a cosmetics cleanroom project. The “class 100,000” label appears without stated particle size, unit, test state, or method; it must not be converted to an ISO class on that basis.
Transferable project lessons
Reported facts. The public case page reports the project size, the use of modular construction across multiple areas, and integration of HVAC and support spaces. Transferable lesson. Large-area modular installation requires coordination across zones and disciplines, and the same coordination discipline applies to cosmetics facilities of comparable scale. Not established. The page does not establish the project’s difficulties, acceptance results, or customer benefits; those are not inferred here.
Explicit limitations
This case is not a cosmetics facility. Food and beverage regulation is not interchangeable with cosmetics GMP. The “Class 100,000” designation should not be converted to an ISO class without knowing the test method, particle size, test state, and reporting basis. The claim is attributed to Deiiang’s case page, not presented as an independent finding. Images from the case page are used only as food/beverage context, with source attribution to the public case page. No additional permission is claimed.
A Procurement Checklist for Cosmetics Manufacturers
Use this checklist to structure the evaluation and avoid gaps in the project definition.
- Product and process risks identified and documented
- Target markets and applicable regulatory requirements confirmed
- Design and utility responsibilities defined between parties
- Environmental criteria (temperature, humidity, pressure, cleanliness where applicable) agreed
- Component data reviewed with clear understanding of scope and test conditions
- Test and validation scope defined, including methods, acceptance criteria, and the party that will execute testing
- Instrument calibration requirements identified
- Defect correction and re-test process agreed
- Spare parts and filter replacement path defined
- Filter-change access route and maintenance clearances confirmed
- Local service and support time zone confirmed
- Change-control re-test conditions agreed
- Training and maintenance responsibilities assigned
- Local installation and after-sales support confirmed
Frequently Asked Questions
Is one ISO class mandatory for every cosmetics facility?
No. There is no universal ISO class requirement for cosmetics manufacturing. The appropriate level of control depends on the product, the process, the risk assessment, and applicable market regulations. See the ISO and GMP layers section for the distinction between classification and GMP compliance.
How does ISO 14644 differ from ISO 22716?
ISO 14644-1 classifies air cleanliness by particle concentration. ISO 22716 provides GMP guidelines for cosmetics production, covering documented controls across the manufacturing operation. They address different aspects of facility and operational control. See the comparison table for a side-by-side view.
What is included in a one-stop solution?
A one-stop solution may include requirements definition, design, component manufacturing, installation, commissioning, and handover documentation. Scope varies by provider and project. Owner quality and regulatory responsibilities remain with the manufacturer. See the project workflow for the typical activity split.
What is tested during validation?
Typical checks include airflow and balancing, pressure differentials, temperature and humidity, airborne particle counts where specified, filter integrity where specified, controls and alarms, and documentation. The protocol is project-specific. See the validation section for the full list.
Can a modular room be expanded or reconfigured?
Modular construction is generally designed to support expansion and reconfiguration more readily than stick-built construction. Feasibility depends on the specific system, available space, utility capacity, and HVAC design. Expansion planning should consider utility headroom and system capacity from the initial design.
Conclusion: One Integrated Project, Multiple Layers of Evidence
A successful modular cleanroom for cosmetics brings together several layers of work: process risk assessment, facility design that follows the process, modular construction and installation, verification against agreed criteria, GMP systems that address the broader quality controls, and lifecycle support for maintenance and change. No single layer is sufficient on its own. A particle classification without GMP systems is incomplete. GMP procedures without appropriate physical controls may not manage contamination effectively.
Deiiang’s product range covers components and systems relevant to these facilities, including modular clean rooms, clean booths, and FFUs. Manufacturer product information describes components and systems; it is not a project certification, and it does not by itself establish that a specific facility meets a given classification or GMP requirement.
To move forward with a specific project, the next step is to gather the relevant inputs: product type and formula characteristics, process flow and open/closed operations, site location and building constraints, target market and regulatory basis, available utilities, and desired schedule. With those inputs, the design and validation scope can be defined on a project-specific basis. No performance, compliance outcome, savings, or delivery timing is promised without project evidence.
References
- ISO 14644-1:2015 — Classification of air cleanliness by particle concentration
- ISO 22716:2007 — Cosmetics — Good Manufacturing Practices (GMP)
- Regulation (EC) No 1223/2009 on cosmetic products, Article 8
- European Commission — Cosmetic products and harmonized standards
- U.S. FDA — Modernization of Cosmetics Regulation Act of 2022 (MoCRA)
- Deiiang — Modular Clean Room
- Deiiang — Clean Booth
- Deiiang — FFU
- Deiiang-reported food/solid-beverage reference project (public case page)